Here, we describe a protocol for the optimization and parameterization of amino acid residues modified with reactive carbonyl species, adaptable to protein systems. The protocol steps include structure design and optimization, charge assignments, parameter construction, and preparation of protein systems.
Method Article
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
In This Article
Summary
Abstract
Protein carbonylation by reactive aldehydes derived from lipid peroxidation leads to cross-linking, oligomerization, and aggregation of proteins, causing intracellular damage, impaired cell functions, and, ultimately, cell death. It has been described in aging and several age-related chronic conditions. However, the basis of structural changes related to the loss of function in protein targets is still not well understood. Hence, a route to the in silico construction of new parameters for amino acids carbonylated with reactive carbonyl species derived from fatty acid oxidation is described. The Michael adducts for Cys, His, and Lys with 4-hydroxy-2-nonenal (HNE), 4-hydroxy-2-hexenal (HHE), and a furan ring form for 4-Oxo-2-nonenal (ONE), were built, while malondialdehyde (MDA) was directly attached to each residue. The protocol describes details for the construction, geometry optimization, assignment of charges, missing bonds, angles, dihedral angles parameters, and its validation for each modified residue structure. As a result, structural effects induced by the carbonylation with these lipid derivatives have been measured by molecular dynamics simulations on different protein systems such as the thioredoxin enzyme, bovine serum albumin and the membrane Zu-5-ankyrin domain employing root-mean-square deviation (RMSD), root mean square fluctuation (RMSF), structural secondary prediction (DSSP) and the solvent-accessible surface area analysis (SASA), among others.
Introduction
In the constant pursuit of understanding the molecular behavior of proteins with oxidative modifications, computational chemistry has become a fundamental pillar in the broad field of scientific research. This relies on the use of theoretical models capable of interpreting physical phenomena in electronic systems, using mathematical equations to describe the atomic behavior of molecules. Within this landscape, computational simulations of proteins stand out as crucial tools to analyze the atomic behavior of molecular systems. Based on the evaluation of structural behavior, energetic calculations, and conformational states1, these methods become strategic allies to predict the behavior of biomolecular systems.
These simulations specialize in studying structural changes and assessing the loss or gain of biological functions in protein systems. However, computational approaches have shown significant limitations when applied to protein systems containing modified residues formed by covalent post-translational modifications in the sequence. This is because many available methods lack resources with parameters adaptable to force fields that are compatible with the most common packages of programs for molecular dynamics simulations of proteins2,3,4,5,6. Therefore, the standardization of computational software-compatible force-field adaptive parameters is essential to facilitate the precise coupling of topologies and atomic coordinates with the equation governing the potential energy of the system7.
In response to these challenges, a protocol adaptable to new modified amino acid residues with aldehydes derived from lipid peroxidation has been developed using ab initio methods. In that sense, the optimization of the structural geometry of the new residues allows the assignment of adaptive charges to new bond, angle, and dihedral parameters that can be run in general force fields such as AMBER. Subsequent validation of these parameters allows the determination of the consistency and robustness of the method applicable to molecular dynamics simulations.
One of the notable strengths of this method lies in its ability to adapt to diverse post-translational modifications, from carbonylation to phosphorylation, acetylation, and methylation, among others. This versatility is not only limited to protein systems but extends to macromolecular structures, allowing coupling with atomic topologies and coordinates. In contrast, previous studies reveal that the standard parameterization of post-translational modifications is only suited to a specific type of modification and can only be obtained from published repositories, lacking the ability to create new structures8.
Currently, challenges in protein structure prediction and design are becoming more evident when modeling structures with post-translational modifications. The scarcity of parameters describing alterations at specific amino acid sites underlines the urgent need to develop and apply computational methods that can be adjusted to standard parameterizations. The aim of this protocol is to provide a route for the in silico construction of new parameters for amino acids covalently modified with reactive carbonyl species derived from fatty acid oxidation. These modified amino acids are recognized by the general amber force field (GAFF) and can, therefore, be used to evaluate in silico the structural and functional effects that this kind of carbonylation has on their target proteins.
Protocol
1. Design and optimization of the new modified amino acid
NOTE: This stage involves drawing the structures of modified residues and optimizing their energy.
- Designing the modified structures and optimizing their structure.
- Use a computational chemistry software package to draw the amino acid molecules bound to the reactive aldehydes derived from lipid peroxidation, i.e. with HNE, HHE, MDA and ONE. Once modified, at the carboxyl group end of the amino acid draw the shape of the methylamine group. At the amino end, draw an acetyl group to emulate the peptide bonds of the modified amino acid, as shown in Figure 1.
- Click on the Clean Icon for structure cleanup. For structure optimization, click on Calculate > Gaussian calculation setup… or Ctrl+G, then click on General and uncheck Write connectivity. Click on Job Type > Optimization, as shown in Figure 2. In additional keywords type the following line:
SCF=tight test Pop=MK iop(6/33=2) iop(6/42=6) opt
NOTE: Here, GaussView automatically sets Hartree Fock (HF) as the functional and the basis set to 3-21. HF is commonly used as a functional in various applications, although other functionals, such as M062X, have also been used, depending on the specific system and the researcher's objectives. Remember that since it is a neutral charge molecule, the charge and multiplicity must be 0 and 1, respectively. - To change the basis set, click on Method > 6-31G for the Basis set.
- To execute the optimization on the same computer, click on Submit. To optimize from a Gaussian terminal, write the following command:
g16 name_of_the_file.com & - Click on File > Save. Save the file as .com for Linux or. gjf for Windows. Once the optimization is complete, open the output file (.out in Windows and .log in Linux) and verify that everything went well. There should be no error messages at the end of the document.
NOTE: If there are no error messages at the end of the output file, it means that the optimization was performed correctly.
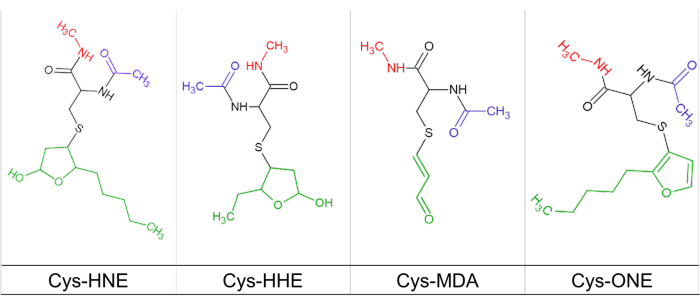
Figure 1: Cysteine modified with reactive carbonyls. Representation of the chemical structure of cysteine (black line) modified with HNE, HHE, MDA and ONE (green line), and linked with acetylamide (blue line) and methylamide (red line) substituent groups. Please click here to view a larger version of this figure.
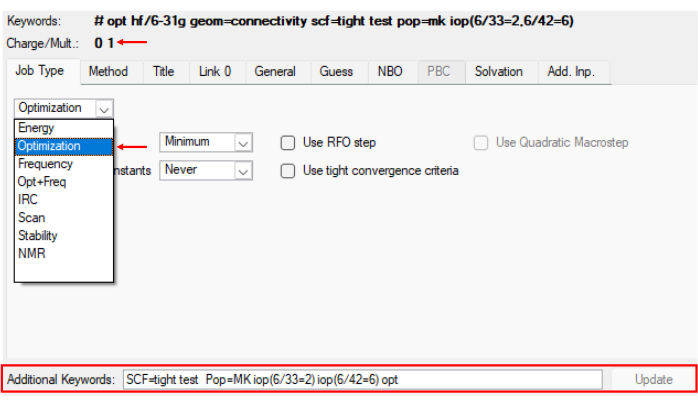
Figure 2: Menu to optimize modified residues synthetized. Reference image illustrating step 1.1 of the protocol, which shows the optimization step of the modified structure in Gaussian program. Please click here to view a larger version of this figure.
2. Parameterization of the modified amino acid residues
- Create the prepin file using the antechamber program from the AmberTools 16 package or the available version. Refer to Figure 3 for an image of how the prepin file should look like.
antechamber -i init-gau.log -fi gout -o u00.prepin -fo prepi -c resp -s 2 -rn U00 -at gaff2 -nc 0
NOTE: Hereafter, the italicized text corresponds to the file name and varies according to the researcher's criteria. In this case, init-gau.log corresponds to the file obtained after optimization. - For building the parameter file, type the following command:
parmchk -i u00.prepin -f prepi -o u00.frcmod
At this point verify that the. frcmod file has been created. Refer to Figure 3 for an example of what the .frcmod file might look like. - Building the library file
- Open XLEaP, universe editor with the xleap command. A window resembling the one shown in Figure 4 will open. Then, follow the steps below to generate the library file that contains relevant data. Type the following commands:
source leaprc.gaff2
loadamberparams u00.frcmod
loadamberprep u00.prepin
list
NOTE: Verify that the U00 file has been created by using the list command. - Edit the ends of modified structures and adjust the resulting charges by typing the following commands:
edit U00
A graphical interface will be displayed (see Figure 4). - Select Erase option. Click on the Atoms of the Acetyl and Methylamine Ends that were added in step 1.1 to delete them (see Figure 4 for a reference of what the carboxyl and amino ends of the modified residue should resemble).
- Charge neutralization
- At this point, the molecule's charge is no longer neutral due to the elimination of atoms in step 2.3.3. The charge comes from both the carboxyl end and the amino end. To neutralize both the charge of the amino end and the carboxyl end, follow the steps below.
- To obtain the total charge value (see Figure 5), type:
charge U00
Divide by two the charge obtained. Use the absolute value for the total charge value. - In the graphical interface select the Whole Molecule. Click on Display > Names. Click on Edit > Edit selected atoms. At this point a window with a table should appear.
- Verify the name of the N and C terminal atoms. In the table, add the value obtained for the division of the total charge (absolute value; see Figure 5). Then save and quit by clicking on Table > Save and quit.
- Ensure the charge is zero (see Figure 5):
charge U00 - To exit the program and save the library file type:
desc U00
saveoff U00 u00.lib
quit - Verify that the library file (.lib) has been created correctly (see Figure 6 for reference).
- Open XLEaP, universe editor with the xleap command. A window resembling the one shown in Figure 4 will open. Then, follow the steps below to generate the library file that contains relevant data. Type the following commands:
- Build the pdb file of the modified residue with the new parameters as described below.
tleap
source leaprc.gaff2
loadamberparams u00.frcmod
loadoff u00.lib
x = U00
savepdb U00 from-lib.pdb
quit - Preparation of the protein
- Download the PDB file of the protein to be modified. Thioredoxin was selected as the model protein system (PDB ID: 2IFQ). Use an appropriate protein visualizer to erase water molecules, dimers (if necessary), ligands, etc.
NOTE: This step can be performed in viewers such as UCSF Chimera or Discovery - Add the file from-lib.pdb (file obtained in step 2.4) and overlay it on the amino acid residue to be modified (as shown in Figure 7). Ensure the amino and carbonyl terminal ends of the from-lib.pdb match the amino acid to be modified.
- Delete the protein, only the from-lib.pdb file should remain in the three-dimensional space occupied by the residue to be modified. Remove H from the N- and C-terminal atoms.
- Save the from-lib.pdb as u00-moved.pdb with the new coordinates.
- Once the coordinates of the modified residue have been saved, with a text editor open the u00-moved.pdb and the protein PDB file that was cleaned previously. Here, we use Notepad++ v8.4.8 text editor.
- Copy the coordinates from u00-moved.pdb as shown in Figure 8 and paste them into the protein pdb file, replacing the residue to be modified. This is intended to adapt the bond between the modified residue and the protein system.
- Adjust the typology to be compatible with the protein PDB format, changing HEATATM to ATOM and change the numbering 1 to the one corresponding to the residue to be modified. Save the new file as complex.pdb.
- Download the PDB file of the protein to be modified. Thioredoxin was selected as the model protein system (PDB ID: 2IFQ). Use an appropriate protein visualizer to erase water molecules, dimers (if necessary), ligands, etc.
- Generation of modified protein-residue binding connections
- In the protein visualizer program open the file from-lib.pdb. Select the entire structure. Click on Structure > Labels > Add... > OK.
- Verify the nomenclature assigned to the N- and C-terminal atoms. In another window, open the u00.lib file in the text editor.
- In the list that appears, verify the position of N- and C-terminal, taking into account the nomenclature assigned.
- In the u00.lib file, locate the line: !entry.U00.unit.connect array int. Below that line, two numbers will appear. Change the first number to the position of the N-terminal and change the second number to the position of the C-terminal and save.
- Create the parameter list by typing the following lines:
tleap
source leaprc.gaff2
source leaprc.protein.ff14SB
loadoff u00.lib
loadamberparams u00.frcmod
x = loadpdb complex.pdb
check x
NOTE: At this point, tleap will provide a list of bonds, angles, and dihedral angles to parameterize. - Typology identification
- Open the complex.pdb file in the protein visualizer. Select the modified residue and the adjacent residues on either side.
NOTE: In the tertiary structure of the protein, it is common for a gap to occur at the site of the modified residue. - Display ball and stick structure for the selected residues. Display nomenclature only for the modified residue as shown in step 2.6.1 Open library file (.lib) in the chosen text editor.
- Based on the nomenclature observed, identify in the library file (.lib) the assigned topology (found in quotation marks next to the nomenclature) that corresponds to the one used in the list of bonds, angles, and dihedral angles to be parameterized, created in step 2.7.
NOTE: In the list of bonds, angles, and dihedral angles provided by tleap, capital letters represent the atoms of the amino acids adjacent to the modified residue.
- Open the complex.pdb file in the protein visualizer. Select the modified residue and the adjacent residues on either side.
- Parameterization of bonds, angles, and dihedral angles with parmcal (a program in Amber)
NOTE: For this step, it will be necessary to use the parmcal program of the Amber package. It will also be indispensable to have the frcmod file (u00.frcmod) and the library file open in the text editor. The protein visualizer should be used to visualize the angles and bond distances. In the protein visualizer, the amino acid residues that are bound to the modified one will be selected in order to generate the bond distances, angles, and dihedrals (see list step 2.7). These data will be implemented to calculate the constants in parmcal and add them in the frcmod file for the parameter's creation.- Generation of bond distances and angles in the Visualizer
- In the Visualizer, select the atoms involved in the bond or angle. Click on Structure > Monitor > Distance or Angle.
- Perform the following procedure for each new parameter to be added. The data to be entered in parmcal is indicated in bold. Below is an example of how to create the binding parameter between the N-terminal of the modified amino acid and the adjacent C of the other amino acid.
Parmcal
Please select:
0. set parameter se (gaff)
1. calculate the bond length parameter: a-b
2. calculate the bond angle parameter: a-b-c
3. exit
0
Please select which parameter set to use: 1-gaff (the default) or 2-gaff2
2
Force field parameters set has been set to gaff2
Please select:
0. set parameter se (gaff)
1. calculate the bond length parameter: a-b
2. calculate the bond angle parameter: a-b-c
3. exit
1
Please input the element name of atom A in A-B
C
Please input the element name of atom B in A-B
ns
Please input the bond length in non-positive number
means to calculate it according to empirical rules
1.455
BOND C-ns 270.256 1.455
NOTE: The double underline is copied and added in the frcmod file. For this example, it is added below the last line of the BOND section. The dihedral angles are added according to the values reported by Alviz-Amador et al.9. - After creating all the bond, angle, and dihedral parameters and adding them to the frcmod file, save the frcmod file, ensuring that the new parameters are included.
- Generation of bond distances and angles in the Visualizer
- For topology and coordinate file generation, type the following commands:
tleap
source leaprc.gaff2
source leaprc.protein.ff14SB
loadoff u00.lib
loadamberparams u00.frcmod
x = loadpdb complejo.pdb
source leaprc.water.tip3p
charge x- Add the number of Na or Cl ions necessary to neutralize the charge by typing:
addions x Na+ 5
solvateoct x TIP3PBOX 10.0
saveamberparm x prot.topo prot.coords
NOTE: If it is desired to add Cl ions instead of Na, replace Na+ by Cl-. The 5 corresponds to the number of ions to be added and is adjusted in order to neutralize the charge.
- Add the number of Na or Cl ions necessary to neutralize the charge by typing:
- For molarity calculation type:
tail -f prot.coords- Copy the final line produced, substituting it with the content in bold within the subsequent instruction. 0.15 corresponds to the target molarity.
usr/bin/perl molarity.perl 0.15 101.3356150 101.3356150 101.3356150 109.4712190 109.4712190 109.4712190 Kconts
This will generate the amount of Cl- and Na+ ions to be added, as described in step 2.10. At this point the generation of the topology and coordinate files of the modified amino acid residue with the new parameters takes place.
- Copy the final line produced, substituting it with the content in bold within the subsequent instruction. 0.15 corresponds to the target molarity.
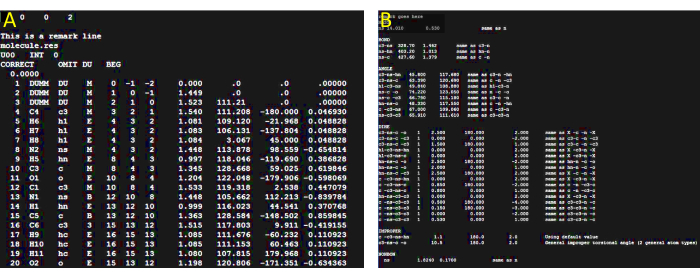
Figure 3: Parameter file preparation. (A) Reference image illustrating the expected appearance of the prepin file generated in step 2.1. The visualization of the file was conducted using the GNU nano text editor v2.3.1. (B) Reference image illustrating the expected appearance of the frcmod file generated in step 2.1. Please click here to view a larger version of this figure.
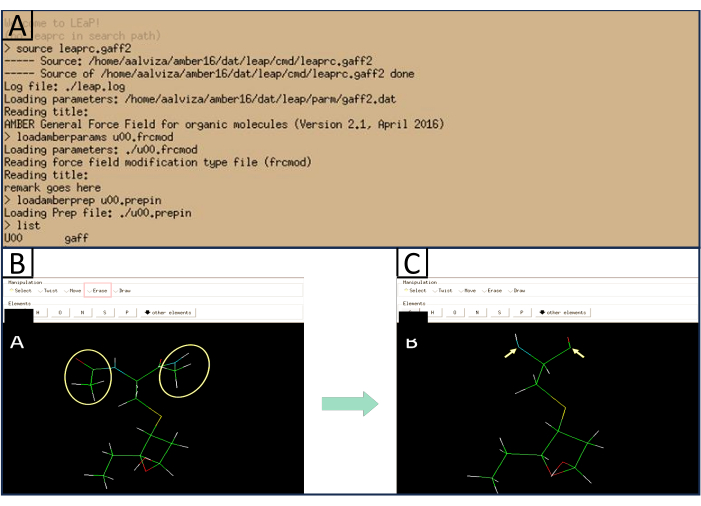
Figure 4: Reference image of XLEaP window. (A) Shows the expected response when typing the mentioned commands.(B) Shows the atoms that need to be removed (yellow) and the option that needs to be selected in order to do so (red). (C) Shows a reference image of what the amino and carbonyl terminal ends of the modified residue should look like after the acetyl and methylamine groups are deleted. Please click here to view a larger version of this figure.
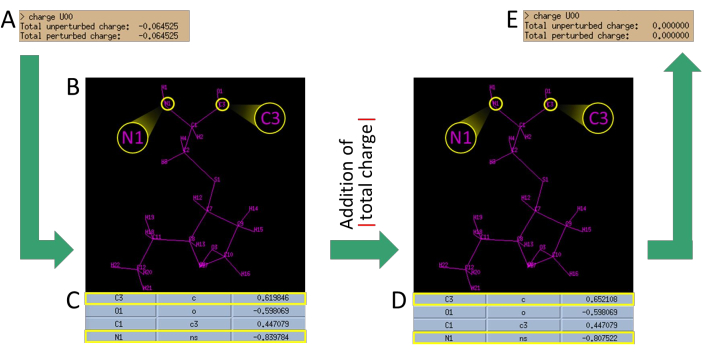
Figure 5: Charge neutralization procedure. (A) Calculation of the total charge after the removal of acetyl and methylamine groups. (B) Determination of the assigned nomenclature for the atoms of the residue. Pay attention to the assigned nomenclature for the N of the amino terminal and C of the carboxyl terminal. (C) Identification of the assigned charges for these two atoms (N1 and C3) in the table. Take the charge value of the atoms (divided by 2) and add the absolute value of the obtained charge. (D) Substitution of the charge values of N1 and C3 with the obtained values. (E) Verification that the resulting charge is now zero. (all data provided is for reference only and may vary depending on the modified residue). Please click here to view a larger version of this figure.
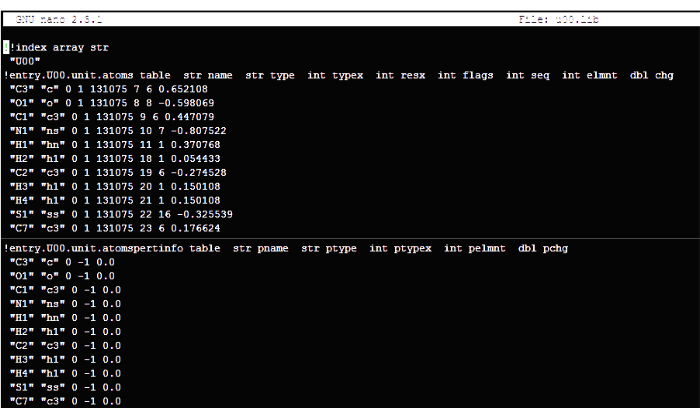
Figure 6: Reference image of the desired structure of the library file (.lib). It is important to note that the image provided only displays a condensed representation of the complete file. Please click here to view a larger version of this figure.
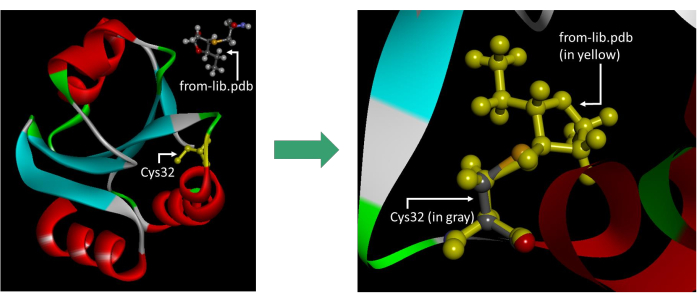
Figure 7: Reference image illustrating the correct positioning of the from-lib.pdb file. It is important to note that the displayed image includes the hydrogens on the N and C termini, which should be excluded before saving the file. The image was taken in the Visualizer software. Please click here to view a larger version of this figure.
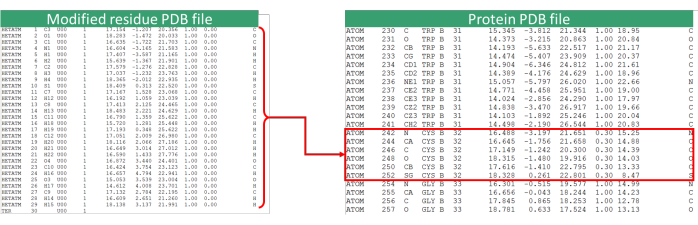
Figure 8: PDB file update. Reference image of the procedure for replacing the residue coordinates (in this case Cys32) with the modified residue. The modified residue PDB file refers to the u00-moved.pdb file. Please click here to view a larger version of this figure.
Results
To illustrate the implementation of the protocol and evaluate the results, the following analyses will be considered. The data set generated by assigning new parameters to modified amino acid residues was constructed based on optimization of the electronic structures, which were supported for partial RESP loadings. Figure 9 shows the structural conformation of one of the amino acid residues optimized with the parameter assignment.
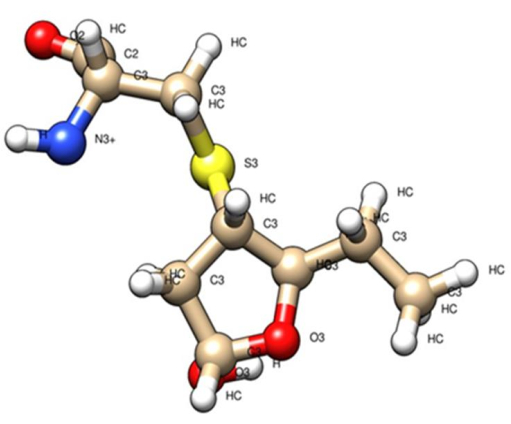
Figure 9: Cys-HHE residue synthesized in silico. Representation of HHE-modified cysteine amino acid with assigned topology and coordinate parameters. Please click here to view a larger version of this figure.
The structures obtained from theoretical DFT levels with M062X/6-31G were compared with the classical mechanics structures through molecular dynamics simulations in AMBER. Each of the parameters obtained from the simulations showed a good correlation with the theoretical data from quantum mechanics. The average bond distance errors showed values of approximately 0.001 - 0.002 Å, while the angles were ~ 8.2°. The typology, distances and constants of bonds and angles are listed in Table 1. These data were similar to those reported in the data article by Alviz-Amador et al.9. Parameter files are available at http://research.bmh.manchester.ac.uk/bryce/amber/.
| Cys-HHE | ||||||
| Methods | Bond | Angle | ||||
| (Å, ± Stdev) | (°, ± Stdev) | |||||
| QM | S1 –C4 | C6-C8 | C8-C9 | S1-C4-C5 | O2-C6-C8 | C6-C8-C9 |
| (m062x/631g(d) | 1.82 | 1.52 | 1.53 | 115.9 | 109.25 | 112.21 |
| MM (AMBER) aa alone | 1.85± | 1.55± | 1.54± | 111.66± | 109.77± | 113.16± |
| 0.002 | 0.002 | 0.002 | 0.152 | 0.14 | 0.148 | |
Table 1: Bond distance and angle parameters comparison. The values of bond distances and angle obtained by quantum (QM) and classical methods showed no significant differences.
Once each of the parameters for the modified amino acid residues was generated and validated, the dynamic behaviors were examined through molecular dynamics simulations with trajectories of 1 µs in order to evaluate the effect on the stability of each residue compared to its native counterpart (Figure 10). The RMSD values obtained for each of the modified amino acids did not show significant differences from their native counterpart, and they maintained their conformational stability throughout the entire trajectory.
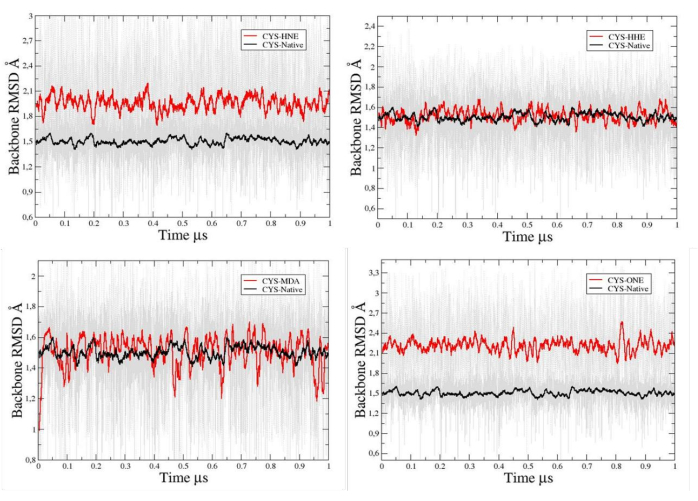
Figure 10: RMSD graph of synthesized residues in silico. Representative RMSD of unmodified and modified cysteine residue with HHE, HNE, MDA and ONE. Please click here to view a larger version of this figure.
The files resulting from the parameterization of modified amino acid residues have been utilized to substitute natural structural amino acids in proteins that have experimental evidence of carbonylation. This substitution was undertaken to evaluate the structural and functional impacts that may occur in the protein as a result of these modifications. It had been reported by in silico studies of carbonylation by reactive carbonyl species on protein systems like Ankyrin and Thioredoxin10,11.
Discussion
One of the critical steps in developing the AMBER parameterization protocol was the quantum optimization of the new amino acid residues modified with the lipid peroxidation derivatives, due to the energetic variability related to the minimization and the way of assigning RESP charges in the AMBER antechamber. For this, ab initio optimization methods with Hartree-Fock (HF/6-31G) and semiempirical density functional theory (DFT; B3LYP/6-31G and M062X/6-31G) were established to evaluate the response to the load assignment. As a result, the HF functional presented better performance/computational cost ratio, taking this into account as a previous step to the protocol. This was also demonstrated in the study by Zhou et al.12.
During the application of the protocol there can be several sources of errors. The possible steric hindrances generated by the additional structures of the modification often lead to errors that are often solved through the minimization steps of the molecular system. On the other hand, the parameters of the dihedral angles are usually adjusted at the end of the parameterization process and therefore, sometimes they tend to show as a possible error, in this particular case it is suggested to adjust the parameters by homology, as reported by Alviz-Amador9 and add it in the new format to eliminate the error.
One of the limitations of the method is the effort required for the step-by-step development of the parameterizations. The generation of new parameters from the new electronic structures and then adapting these parameters to protein systems requires a lot of dedication for its good execution. Therefore, a good strategy when implementing our protocol is to follow the step-by-step instructions and read the guide carefully.
In the landscape of molecular dynamics simulations, the significance of the AMBER protocol becomes evident. Its adaptive nature and versatility make it a valuable tool for researchers exploring diverse research areas. Beyond its application in protein systems, its extension to macromolecular structures opens doors to new possibilities. This adaptability not only addresses the existing gaps in standard parameterization methods but also offers a pathway for the creation of novel structures, thus expanding the horizons of molecular dynamics research. On the contrary, other investigations demonstrate that the conventional parameterization of post-translational modifications is limited to a particular modification type and is exclusively derived from publicly available repositories8, lacking the capability to generate novel structures.
Modifications resulting from the presence of reactive carbonyl species are frequently associated with a range of pathologies, including cancer, metabolic disorders, and degenerative diseases following different mechanisms13,14 . The support provided by this protocol is useful to assess various crucial properties, such as conformational stability, atomic flexibility, loss of secondary structures, solvent accessibility, and protein-protein interaction energy, among others. Consequently, the measurement of these properties could prove beneficial in situations where carbonylated proteins can induce irreversible alterations in biological systems, leading to conformational instability, increased or decreased atomic flexibility, and loss of secondary structure10,11.
In conclusion, the AMBER parameterization protocol, with its critical steps, adaptability, and versatility, stands as a pioneering method in the realm of molecular dynamics simulations. While acknowledging its limitations, its significance is underscored by its ability to address the shortcomings of existing methods, providing researchers with a powerful tool to explore the intricacies of molecular structures and behaviors across a spectrum of biological and chemical systems.
Disclosures
The authors have nothing to disclose.
Acknowledgements
This work was supported by research grant code 1107-844-67943 from Ministerio de Ciencia, Tecnología e Innovación (Minciencias) and the University of Cartagena (Colombia) for grant to support the research groups 2021 and Acta 017-2022.
Materials
| Name | Company | Catalog Number | Comments |
|---|---|---|---|
| AmberTools16 or Upper | The Amber Project | Amber is a suite of biomolecular simulation programs | |
| Gaussian 09 or Upper | Gaussian Inc | Draw and optimize structures | |
| Linux Ubuntu | GNU/Linux | Platform for AmberTools | |
| NVIDIA GPUs GTX 1080 or Upper | Nvidia | Compatible with PMEMD |
References
- Cornell, W. D., et al. A second generation force field for the simulation of proteins, nucleic acids, and organic molecules. J Am Chem Soc. 117 (19), 5179-5197 (1995).
- Wang, J., Wolf, R. M., Caldwell, J. W., Kollman, P. A., Case, D. A. Development and testing of a general amber force field. J Comput Chem. 25 (9), 1157-1174 (2004).
- Brooks, B. R., et al. CHARMM: A program for macromolecular energy, minimization, and dynamics calculations. J Comput Chem. 4 (2), 187-217 (1983).
- Mayo, S. L., Olafson, B. D., Goddard, W. A. DREIDING: a generic force field for molecular simulations. J Phys Chem. 94 (26), 8897-8909 (1990).
- Daura, X., Mark, A. E., van Gunsteren, W. F. Parametrization of aliphatic CHn united atoms of GROMOS96 force field. J Comput Chem. 19 (5), 535-547 (1998).
- Robertson, M. J., Tirado-Rives, J., Jorgensen, W. L. Improved peptide and protein torsional energetics with the OPLS-AA force field. J Chem Theory Comput. 11 (7), 3499-3509 (2015).
- Guvench, O., MacKerell, A. D. Comparison of protein force fields for molecular dynamics simulations. Methods Mol Biol. 443, 63-88 (2008).
- Petrov, D., Margreitter, C., Grandits, M., Oostenbrink, C., Zagrovic, B. A systematic framework for molecular dynamics simulations of protein post-translational modifications. PLoS Comput Biol. 9 (7), e1003154(2013).
- Alviz-Amador, A., et al. Development and benchmark to obtain AMBER parameters dataset for non-standard amino acids modified with 4-hydroxy-2-nonenal. Data Brief. 21, 2581-2589 (2018).
- Pineda-Alemán, R., et al. Cysteine carbonylation with reactive carbonyl species from lipid peroxidation induce local structural changes on thioredoxin active site. J Mol Graph Model. 124, 108533(2023).
- Alviz-Amador, A., et al. Effect of 4-HNE modification on ZU5-ANK domain and the formation of their complex with β-Spectrin: A molecular dynamics simulation study. J Chem Info Model. 60 (2), 805-820 (2020).
- Zhou, A., Schauperl, M., Nerenberg, P. S. Benchmarking electronic structure methods for accurate fixed-charge electrostatic models. J Chem Info Model. 60 (1), 249-258 (2020).
- Gęgotek, A., Skrzydlewska, E. Biological effect of protein modifications by lipid peroxidation products. Che Phys Lipids. 221, 46-52 (2019).
- Moldogazieva, N. T., Zavadskiy, S. P., Astakhov, D. V., Terentiev, A. A. Lipid peroxidation: Reactive carbonyl species, protein/DNA adducts, and signaling switches in oxidative stress and cancer. Biochem Biophys Res Comm. 687, 149167(2023).
Reprints and Permissions
