Here we present a simplified one-pot platform for the synthesis of deoxyribonucleoside 5′-triphosphates (dNTPs) directly from unprotected deoxyribonucleosides. The protocol eliminates protecting-group strategies and intermediate isolation steps, thereby reducing the number of synthetic operations and lowering the technical barrier for dNTP production in research laboratories.
方法文章
A Simplified One-Pot Platform for the Synthesis of Deoxyribonucleoside 5′-Triphosphates from Unprotected Nucleosides
本文内容
摘要
摘要
Deoxyribonucleoside 5'-triphosphates (dNTPs) are essential biomolecules with broad applications in molecular biology, biotechnology, and pharmaceutical research. Their chemical synthesis, however, is often complicated by the formation of complex mixtures of phosphorylated byproducts, resulting in reduced yields and demanding purification procedures. This limitation has traditionally been addressed using protecting groups to improve regioselectivity and prevent undesired phosphorylation at hydroxyl groups other than the 5′ position. However, protection and deprotection strategies add synthetic steps, cost, and operational complexity to the overall process.
This protocol describes a simplified one-pot platform for the chemical synthesis of deoxyribonucleoside 5′-triphosphates directly from unprotected deoxyribonucleosides. The procedure is based on a modified Ludwig approach employing phosphorus oxychloride and bis(tributylammonium) pyrophosphate as the key phosphorylating reagents, without requiring prior hydroxyl protection or isolation of monophosphate or diphosphate intermediates. The synthesis of deoxyadenosine 5′-triphosphate (dATP) from unprotected 2′-deoxyadenosine is presented as a representative example of the complete workflow. The procedure involves sequential reagent addition in a single reaction vessel, followed by liquid–liquid extraction and reversed-phase chromatographic analysis.
A key feature of this method is its operational simplicity and compatibility with unprotected deoxyribonucleoside substrates. Using dATP synthesis as a proof of concept, the method achieved a chromatographic yield of 67% under controlled anhydrous conditions. The same reaction platform may be adapted to other canonical deoxyribonucleosides following substrate-specific optimization of reagent stoichiometry, reaction conditions, and chromatographic analysis parameters. The protocol requires minimal specialized equipment and basic anhydrous handling techniques, thereby reducing the technical barriers associated with laboratory-scale dNTP synthesis.
引言
Nucleoside triphosphates (NTPs) are fundamental building blocks of nucleic acid metabolism, serving as substrates for DNA and RNA polymerases, as energy carriers, and as key reagents in molecular biology applications ranging from polymerase chain reaction to in vitro transcription1. Among these molecules, deoxyribonucleoside 5′-triphosphates (dNTPs) are indispensable for DNA synthesis, sequencing, amplification, diagnostic assays, and the preparation of modified nucleic acids. Despite their central role, the efficient production of these biomolecules at the laboratory scale remains challenging. Available synthetic routes must balance regioselectivity, conversion efficiency, product purity, reagent cost, and operational complexity. Chemical and enzymatic strategies have been developed to address these challenges, each presenting distinct advantages and limitations.
Among chemical approaches, one of the most widely adopted methods is the synthesis of nucleoside 5'-triphosphates described by Ludwig in 19812. This method treats unprotected nucleosides (Figure 1A) with phosphoryl chloride (POCl₃) in trimethyl phosphate ([MeO]₃PO) as the reaction medium to generate a reactive nucleoside 5’-dichlorophosphate intermediate (Figure 1B). Subsequent treatment of this intermediate with tetraalkylammonium pyrophosphate salts in triethylammonium bicarbonate (TEAB) buffer results in hydrolysis to yield the corresponding nucleoside di- or triphosphate (NDP or NTP, respectively) as the final product2 (Figure 1). The principal advantage of this route is the direct use of unprotected starting materials, eliminating protection and deprotection steps and substantially shortening the synthetic sequence. However, the reaction is highly sensitive to moisture, temperature, reagent stoichiometry, and substrate solubility. Inadequate control of these parameters can promote hydrolysis, incomplete conversion, and the formation of mono-, di-, or other phosphorylated byproducts.
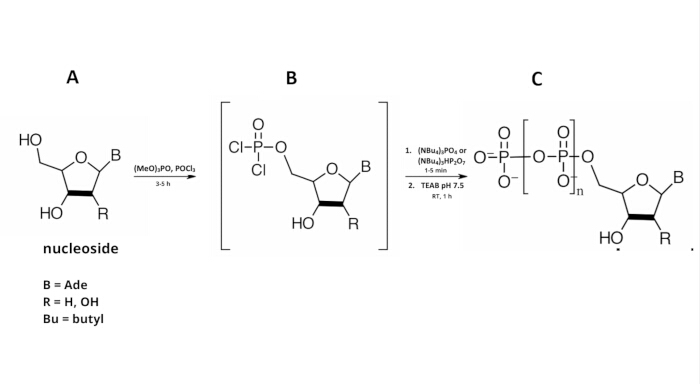
Figure 1: Synthesis of 5'-di- and triphosphate nucleosides via a dichlorophosphate intermediate. (A) Unprotected nucleoside starting material. (B) Reaction with phosphoryl chloride (POCl₃) in trimethyl phosphate ([MeO]₃PO) (3–5 h) generates a reactive 5'-dichlorophosphate intermediate. (C) Subsequent treatment with tributylammonium pyrophosphate or tributylammonium hydrogen pyrophosphate (1–5 min), followed by hydrolysis in triethylammonium bicarbonate (TEAB) buffer at pH 7.5 (room temperature, 1 h), yields the corresponding nucleoside diphosphate (NDP, n = 1) or triphosphate (NTP, n = 2). Please click here to view a larger version of this figure.
In the late 1980s, Ludwig and Eckstein proposed an alternative synthetic route aimed at improving reaction selectivity3,4. This methodology relies on prior protection of the nucleoside, followed by reaction with 2-chloro-4H-1,3,2-benzodioxaphosphorin-4-one (salicyl chlorophosphite) (Figure 2A) to form a cyclic nucleoside phosphite intermediate (Figure 2B). Reaction of this intermediate with tributylammonium pyrophosphate produces a mixture of P(III)-P(V) "trimeta" anhydride species (Figure 2C), which are subsequently oxidized using an iodine/water mixture or elemental sulfur (Figure 2D) to yield a cyclic nucleoside 5'-triphosphate. Acidic or basic hydrolysis, depending on the protecting group employed, then affords the deprotected nucleoside triphosphate as the final product (Figure 2E)4. While this approach offers improved selectivity relative to the direct phosphorylation route, it requires additional protection, oxidation, and deprotection steps that increase both the complexity and the overall cost of the synthesis.
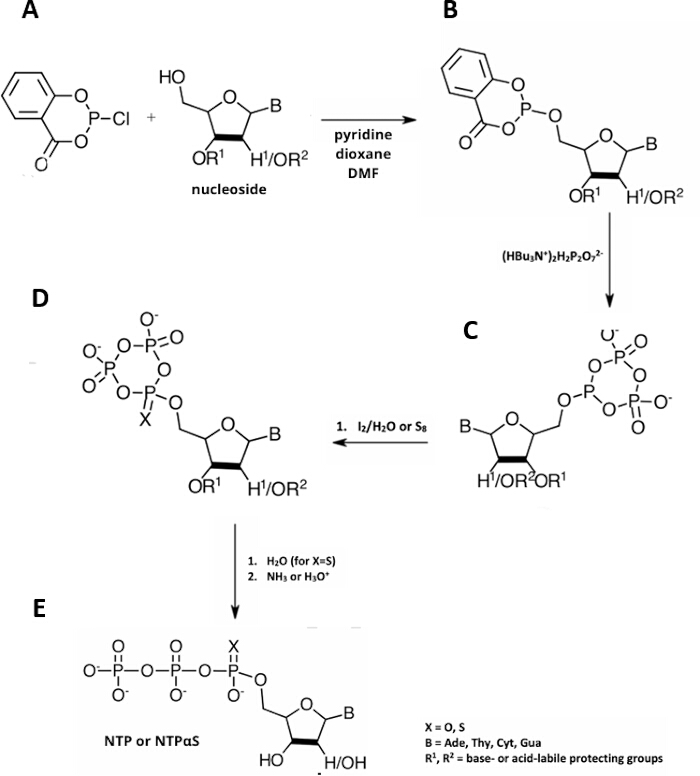
Figure 2: Synthesis of NTP and NTPαS Using the Ludwig and Eckstein method4. (A) Nucleoside protected at the 2'/3'-hydroxyl positions (R1, R2) reacts with salicyl chlorophosphite in pyridine/dioxane/DMF to form (B) a cyclic nucleoside phosphite intermediate. (C) Reaction with tributylammonium pyrophosphate or a thiopyrophosphate analog yields a P(III)–P(V) "trimeta" anhydride intermediate, which is (D) oxidized using either I₂/H₂O (X = O) or elemental sulfur (X = S, for NTPαS synthesis). (E) Acidic or basic hydrolysis of the protecting groups (dependent on their nature) affords the final nucleoside triphosphate (NTP) or phosphorothioate triphosphate (NTPαS) product. Please click here to view a larger version of this figure.
The methodologies established by Ludwig and Eckstein laid the groundwork for the subsequent development of related chemical synthesis routes targeting similar nucleotide products. Mohamady and Taylor5, for example, described a novel strategy for the synthesis of dinucleoside pentaphosphates (Np5N) and nucleoside tetraphosphates (Np4) using trimetaphosphate (TriMP) as the phosphorylating agent in place of phosphoryl chloride. In this approach, the TriMP salt is reacted with mesitylenesulfonyl chloride (MstCl) in the presence of N-methylimidazole (NMI) in dimethylformamide (DMF), followed by the addition of a nucleoside monophosphate (NMP), as outlined in Figure 3. Initial efforts to apply this strategy directly to unphosphorylated nucleosides yielded little to no product; to overcome this limitation, Mohamady and Taylor5 proposed varying the stoichiometric ratio of reagents and optimizing the solvent mixture composition to favor productive interaction between the phosphorylating agent and the nucleoside 5'-hydroxyl group.
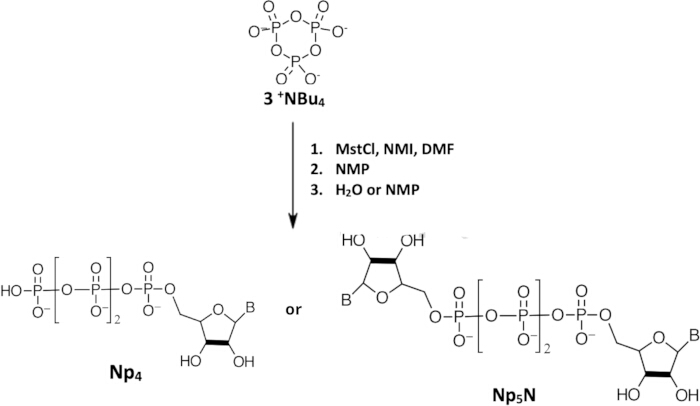
Figure 3: Synthesis of Np4 and Np5N Using TriMP as a phosphorylating agent5. The tetrabutylammonium salt of TriMP (1) is activated with mesitylenesulfonyl chloride (MstCl) and N-methylimidazole (NMI) in DMF, followed by reaction with a nucleoside monophosphate (NMP) and hydrolysis with H₂O or NMP, yielding either a nucleoside tetraphosphate (Np4) or, when a second nucleoside monophosphate is incorporated, a dinucleoside pentaphosphate (Np5N). Please click here to view a larger version of this figure.
Collectively, these studies illustrate that the central challenge in nucleotide chemical synthesis remains achieving an optimal balance between reaction efficiency and cost-effectiveness, particularly when unprotected nucleosides are used as starting materials. In this context, the present protocol describes a simplified one-pot platform for the direct synthesis of deoxyribonucleoside 5′-triphosphates from unprotected deoxyribonucleosides. The workflow is illustrated using the conversion of 2′-deoxyadenosine to deoxyadenosine 5′-triphosphate (dATP) as a representative example. The method employs POCl₃ and bis(tributylammonium) pyrophosphate in a continuous reaction sequence, without isolating monophosphate or diphosphate intermediates. Particular emphasis is placed on nucleoside drying, maintenance of an inert atmosphere, temperature control, reagent stoichiometry, extraction, and chromatographic evaluation, because these parameters are critical for reproducibility. Although substrate-specific optimization may be required, the platform provides a modular basis for adapting the reaction to other canonical deoxyribonucleosides. This protocol is particularly suitable for research laboratories seeking a straightforward route for producing individual dNTPs for biochemical assays, nucleic acid synthesis, or downstream workflows involving chemically or biologically derived nucleoside precursors.
方案
1. Preparation of reagents
NOTE: Although each reagent used in this protocol is commercially available, in-house synthesis of some was performed to reduce costs and improve availability. Both reagents were prepared once in bulk and stored for use across multiple reaction batches.
- Synthesis of triethylammonium bicarbonate (TEAB).
- Add 375.67 mL of triethylamine to 1 L of Type I water under constant stirring to prepare a 2 M solution. Confirm that the initial pH of the solution is approximately 12 using pH strips.
- Generate CO₂ gas by gradually adding dry ice to an Erlenmeyer flask containing water, connected to the triethylamine solution, so that the released CO₂ gas is bubbled into the reaction flask.
NOTE: If gas evolution is insufficient to drive the reaction, add CO₂ pellets directly to the reaction solution to accelerate the process and ensure adequate CO₂ availability. - Continue CO₂ bubbling until the solution reaches a pH of 8–9, as monitored with pH strips, indicating complete conversion to triethylammonium bicarbonate.
CAUTION: Dry ice causes severe cold burns upon direct skin contact; handle using insulated gloves. Perform this step in a well-ventilated area or fume hood to prevent CO₂ accumulation.
- Synthesis of bis(tributylammonium) pyrophosphate.
- Weigh 4.44 g of pyrophosphoric acid and add it to a previously weighed, dry round-bottom flask.
- Add 20 mL of anhydrous acetonitrile (ACN) to the flask.
- Under argon atmosphere, heat the mixture to 50 °C with gentle stirring for 40 min.
- Rotary evaporate the mixture at 40 °C and 300 mbar for approximately 40 min.
- Dissolve the resulting residue in 20 mL of anhydrous ACN under argon atmosphere and heat to 40–50 °C with gentle stirring for approximately 20 min.
- Slowly add 11.91 mL of tributylamine over 5–10 min.
CAUTION: Tributylamine is a flammable and corrosive amine; handle in a fume hood and avoid skin contact. Heat the mixture to 40–50 °C with gentle stirring for approximately 20 min. - Disassemble the system and store the product under argon atmosphere at −20 °C.
NOTE: The estimated concentration of the prepared bis(tributylammonium) pyrophosphate solution is approximately 0.5 M. For a more accurate determination of the final concentration, it is recommended to weigh the reaction vessel at each preparation step to account for changes in mass during solvent removal and reagent addition.
2. Nucleoside preparation
- Weigh 20 mg of 2′-deoxyadenosine (0.08 mmol, 1 eq) and transfer it to a clean, dry reaction flask equipped with a magnetic stir bar.
NOTE: The amount of 2′-deoxyadenosine (dA) used here serves as the reference point for calculating the stoichiometry of all subsequent reagents in this protocol, therefore 1 equivalent equals 0.08 mmol. - Connect the reaction flask to the vacuum and inert gas lines, seal the system, and purge it with argon to establish an inert atmosphere.
- Heat the sealed reaction flask at 80 °C under vacuum for 1 h 30 min to remove residual moisture.
NOTE: If necessary, the procedure could be paused at this point. Store the dried nucleoside under an argon atmosphere until ready to proceed.
3. Nucleoside dissolution
- Under argon atmosphere, add 500 µL of anhydrous trimethyl phosphate (Me₃PO₄) into the sealed reaction system.
- Heat the reaction mixture to 80 °C and stir at 800 rpm for 30 min.
- Add 2 equivalents of tributylamine, calculated relative to the initial amount of nucleoside, and continue stirring at 80 °C for an additional 15 min or until the nucleoside is completely dissolved.
CAUTION: Tributylamine (Bu₃N) is a flammable and corrosive amine; handle in a fume hood and avoid skin contact.
NOTE: For nucleosides with limited solubility in Me₃PO₄, such as 2’-deoxyguanosine, the reaction may proceed as a heterogeneous suspension. In this case, maintain vigorous stirring throughout all subsequent steps.
4. Addition of the phosphorylating agent
- Cool the reaction system to -15 °C while maintaining the argon atmosphere and constant stirring.
- Add 5.8 equivalents of POCl₃ dropwise to the cooled reaction mixture.
CAUTION: Phosphoryl chloride (POCl₃) is a highly reactive, corrosive reagent that releases toxic hydrogen chloride and phosphoric acid vapors upon contact with moisture; handle exclusively in a fume hood with appropriate personal protective equipment (PPE), including nitrile gloves, safety goggles, and a lab coat. - Allow the reaction to proceed for 3 h 45 min at -15 °C under argon atmosphere and constant stirring.
5. Addition of the bis(tributylammonium) pyrophosphate salt
- Pre-cool a mixture of 5 equivalents of bis(tributylammonium) pyrophosphate solution (0.5 M) and 6 equivalents of Bu₃N.
- Under argon atmosphere and constant stirring, add the pre-cooled pyrophosphate mixture dropwise to the reaction mixture at -15 °C.
- Continue stirring the reaction mixture at -15 °C for 4 h under argon.
6. Addition of the quenching agent
- Vent the argon atmosphere and allow the reaction mixture to reach 20 °C.
- With constant stirring, slowly add 2 mL of triethylammonium bicarbonate solution (TEAB, 1 M) to the reaction mixture.
- Stir the quenched reaction mixture at 20 °C for 2 h.
7. Extraction of the reaction crude
- Add 3 mL of Type I water to the crude reaction mixture.
- Transfer the mixture to a separatory funnel and perform three successive liquid-liquid extractions with 5 mL of dichloromethane (DCM) each. CAUTION: DCM is a volatile organic compound and a suspected carcinogen; handle in a fume hood and avoid inhalation or prolonged skin contact.
- After each extraction, discard the organic phase. Retain the combined aqueous phases, which contain the crude nucleoside triphosphate product.
8. Chromatographic analysis of the extracted crude reaction
NOTE: Characterization of the obtained nucleotides was performed by high-performance liquid chromatography (HPLC) using a gradient elution system between mobile phase A (0.1 M KH₂PO₄, pH 6.0) and mobile phase B (0.1 M KH₂PO₄, pH 6.0, 10% v/v CH₃OH). Separation was carried out on a C18 reversed-phase HPLC column (150 mm × 4.6 mm, 5 µm). Detection was performed at 254 nm.
- Prepare a sample vial by dissolving the crude reaction product in the mobile phase A at a 1:100 dilution ratio.
- Set the column temperature to 28 °C and the flow rate to 0.6 mL/min.
- Run the chromatographic separation using the following gradient program over a total run time of 31 min: apply a linear gradient from 0% to 90% mobile phase B over 0–24 min; apply a linear gradient from 90% to 0% mobile phase B over 24–28 min; and hold at 100% mobile phase A from 28–31 min.
NOTE: Ensure the column and mobile phases are equilibrated at 28 °C prior to sample injection to guarantee reproducible retention times between runs. - Determine product yield from the chromatographic data as follows: calculate the peak area under the curve for the product of interest using the integration tool of the LabSolutions software.
- Determine the concentration of the product in the crude reaction extract by direct proportional comparison with the peak area of a commercial standard of known concentration, analyzed under identical chromatographic conditions.
- Use this concentration value to calculate the reaction yield relative to the theoretical yield based on the initial amount of nucleoside used.
结果
The performance of the proposed platform was evaluated using the synthesis of deoxyadenosine 5′-triphosphate (dATP) from unprotected 2′-deoxyadenosine as a representative example. The resulting chromatographic profile illustrates the expected outcome of the protocol and serves as a reference for assessing reaction performance and product formation.
RP-HPLC analysis of the crude reaction mixture (Figure 4) showed that dATP was the predominant reaction product, corresponding to a chromatographic yield of 67%, as determined by peak area normalization against a commercial dATP reference standard (Supplementary File 1). Product yield was determined from the peak area under the curve obtained by chromatographic analysis, using the integration tool of the LabSolutions software. The concentration of the product in the crude reaction extract (Supplementary File 2) was calculated by direct proportional comparison with the peak area of a commercial standard of known concentration, analyzed under the same chromatographic conditions, from which the reaction yield was subsequently determined. Comparison with the commercial standard confirmed the identity of the main product based on its retention time. Furthermore, analysis of the reaction mixture demonstrated complete consumption of the starting 2′-deoxyadenosine, indicating efficient substrate conversion under the optimized reaction conditions.
Besides dATP, several minor peaks were detected in the chromatogram, consistent with the formation of phosphorylated byproducts, including deoxynucleoside mono- and diphosphates, as well as other partially characterized phosphorylated species. The presence of these compounds is commonly observed in one-pot phosphorylation reactions and reflects the competing phosphorylation pathways that occur during nucleotide synthesis. Although the chromatogram corresponds specifically to dATP synthesis, adaptation of the method to the remaining canonical deoxyribonucleosides may require optimization of substrate-specific reaction parameters and chromatographic conditions.
The stoichiometric relationship between phosphorus oxychloride and bis(tributylammonium) pyrophosphate is one of the critical parameters governing reaction performance. Under the optimized conditions described here, the amount of pyrophosphate should remain between 0.3 and 1.0 equivalents lower than that of POCl₃. Increasing the pyrophosphate concentration beyond this range favors the formation of unreacted nucleoside and lower phosphorylated products instead of the desired triphosphate (Figure 5 and Supplementary File 3). Therefore, it is important that the amount of pyrophosphate remains between 0.3 and 1.0 equivalents lower than that of POCl₃.
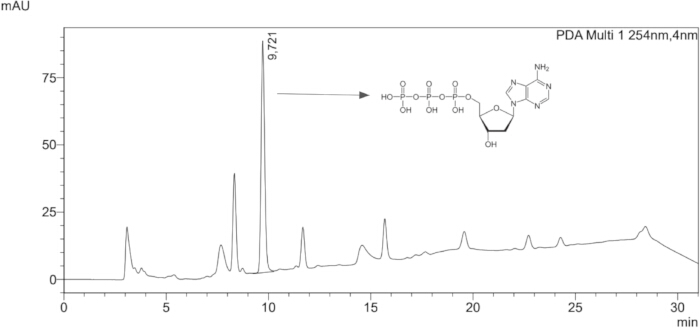
Figure 4: RP-HPLC chromatogram of the crude reaction mixture obtained during the synthesis of dATP. Reversed-phase HPLC analysis (UV detection at 254 nm) of the crude reaction product under optimized stoichiometric conditions, showing dATP as the predominant product. Peak identity was confirmed by co-elution with a commercial dATP standard. Please click here to view a larger version of this figure.
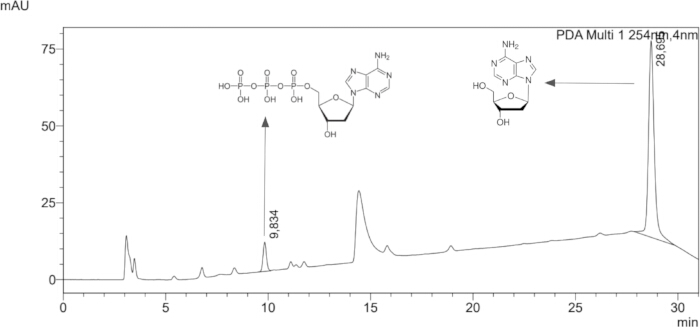
Figure 5: Effect of pyrophosphate stoichiometry on the chromatographic profile of the reaction mixture. RP-HPLC chromatogram of the crude reaction product obtained when the amount of bis(tributylammonium) pyrophosphate exceeded the recommended range of equivalents relative to POCl₃ in the reaction. The unreacted 2'-deoxyadenosine precursor is observed as the dominant peak (retention time ~28.7 min), while dATP is detected only as a minor product, illustrating the adverse effect of pyrophosphate excess on reaction selectivity. The chromatogram corresponds to a PPi:POCl₃ ratio of 1.2:1, with all other reaction parameters maintained as described in the protocol. Please click here to view a larger version of this figure.
Supplementary File 1: Standard dATP 20 ppm chromatogram. Please click here to download this file.
Supplementary File 2: Raw chromatogram of the crude product. Please click here to download this file.
Supplementary File 3: Raw chromatogram of crude reaction product obtained when the amount of bis(tributylammonium) pyrophosphate exceeded the recommended range of equivalents relative to POCl₃ in the reaction. Please click here to download this file.
讨论
Successful implementation of this protocol depends primarily on rigorous control of moisture throughout the reaction. Residual water rapidly hydrolyzes phosphorus oxychloride before it reacts with the nucleoside substrate and may also promote degradation of phosphorylated intermediates and the final triphosphate product6,7. Consequently, complete drying of the nucleoside (step 2.3), the use of anhydrous reagents, and maintenance of an inert argon atmosphere throughout the phosphorylation reaction are essential to maximize product formation and ensure reproducibility. Exposure of the dried nucleoside or reaction mixture to ambient moisture should therefore be minimized during reagent transfer and sampling.
Reaction temperature and reagent stoichiometry are additional parameters that critically influence reaction performance. Maintaining the phosphorylation and pyrophosphate coupling reactions at -15 °C favors selective phosphorylation at the 5′-hydroxyl group while minimizing the formation of undesired phosphorylated products and cyclic polyphosphate species, including trimetaphosphate (Figure 6). Likewise, the molar ratio between phosphorus oxychloride and bis(tributylammonium) pyrophosphate should be carefully controlled, since excessive pyrophosphate favors incomplete phosphorylation, whereas excessive phosphorylating reagent may increase the formation of secondary phosphorylated products. For this reason, optimization of reagent stoichiometry may be required when extending the protocol to additional deoxyribonucleosides.
Efficient stirring is also important, particularly when the nucleoside is only partially soluble in trimethyl phosphate. In such cases, the reaction may proceed as a heterogeneous suspension, and insufficient mixing may limit contact between the substrate and the phosphorylating reagent. The temperature should be allowed to stabilize before each reagent addition, and the addition of POCl₃ and the pyrophosphate solution should be performed slowly to avoid local increases in reagent concentration and temperature.
Although the synthesis of dATP is presented here as a representative example, the methodology should be viewed as a general phosphorylation platform rather than as a compound-specific protocol. The overall reaction sequence—including nucleoside drying, phosphorylation, pyrophosphate coupling, quenching, extraction, and chromatographic analysis—can be applied to other deoxyribonucleosides after appropriate optimization of substrate-dependent parameters, particularly reagent equivalents and chromatographic conditions. Differences in nucleobase reactivity, substrate solubility, and chromatographic behavior should be considered when adapting the procedure to dCTP, dGTP, or dTTP synthesis. Therefore, the conditions reported for dATP provide a starting framework rather than universally optimized conditions for every substrate.
One of the principal advantages of this platform is that it avoids protecting-group chemistry. Conventional synthetic routes frequently require sequential protection, activation, oxidation, and deprotection steps before the desired triphosphate can be obtained, increasing reagent consumption, reaction time, and cumulative material losses. By contrast, the present protocol converts commercially available unprotected deoxyribonucleosides into their corresponding triphosphates through a continuous one-pot reaction sequence, reducing both operational complexity and overall synthesis time. The absence of intermediate isolation steps also decreases the number of handling operations during which product loss may occur.
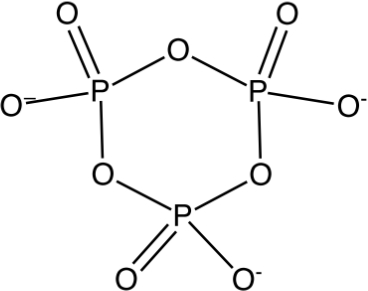
Figure 6: Trimetaphosphate structure, a common byproduct in nucleotide synthesis. Cyclic structure of the trimetaphosphate anion, formed as an undesired polyphosphate byproduct when pyrophosphate is present in excess during the phosphorylation reaction. Please click here to view a larger version of this figure.
Nevertheless, a principal limitation of the described method is the inherent generation of phosphorylated byproducts. The crude reaction mixture may contain deoxyribonucleoside mono- and diphosphates, unreacted precursor, and other incompletely characterized phosphorylated species. Because some of these compounds exhibit similar polarity and chromatographic retention to the target triphosphate, their separation by standard reversed-phase chromatography may be challenging. Further purification is therefore required for applications demanding high-purity dNTPs, adding procedural complexity and potentially reducing the isolated yield. Additionally, the protocol employs moisture-sensitive and corrosive reagents that require careful handling under rigorously anhydrous and inert conditions. Although the procedure does not require highly specialized synthetic equipment, it does require access to an inert-gas or vacuum manifold, efficient temperature control, and personnel trained in handling POCl₃ and volatile organic solvents. These requirements may represent a practical barrier for laboratories without routine experience in anhydrous synthesis.
Preliminary trials applying this protocol to ribonucleosides were performed; however, these remain limited in the number of replicates and have so far yielded low conversion percentages. While the underlying phosphorylation chemistry is, in principle, transferable to ribonucleosides, the additional 2'-hydroxyl group present in ribose introduces a further site of reactivity toward POCl₃ that is absent in 2'-deoxyribonucleosides, favoring a greater diversity and/or proportion of byproducts. Further optimization of reaction conditions is required to establish the applicability and efficiency of this protocol for ribonucleoside triphosphate synthesis, and this remains a priority for future work.
Despite these limitations, the phosphorylation strategy employed here represents a significant practical advantage over multi-step protecting-group-based approaches to nucleoside triphosphate synthesis3,5. Unlike methods that require prior protection of the nucleobase and/or sugar hydroxyl groups followed by selective 5'-activation and subsequent deprotection, the present protocol achieves triphosphorylation directly from the unprotected nucleoside in a single, continuous reaction sequence. This substantially reduces the number of synthetic operations, reagent costs, and total reaction time.
This advantage is consistent with broader trends reported in the nucleotide synthesis literature, where protection–deprotection strategies generally result in lower overall yields compared to one-pot approaches such as the Ludwig and Ludwig-Eckstein methods, owing primarily to the increased number and complexity of the synthetic steps involved. These routes typically require additional operations such as tritylation, acetylation, and a final deprotection step8, each of which introduces a potential loss of material and consequently reduces the cumulative yield of the final compound9. Such multi-step protecting-group sequences have accordingly been described as laborious and time-consuming6. This limitation is particularly pronounced in iterative phosphorylation strategies, some of which require as many as nine individual functional group transformations, substantially complicating scale-up and reducing overall process efficiency10.
Nevertheless, protecting groups may still offer important advantages. Sugar-protecting groups, including ketal-type groups, can accelerate specific transformations and improve chemoselectivity, thereby reducing nucleobase phosphorylation and other undesired reactions that are more difficult to control in direct, unprotected approaches11. Accordingly, protected and unprotected strategies should not be considered interchangeable; rather, each is suited to different experimental priorities.
Taken together, these observations indicate that the choice of synthetic methodology is largely dictated by the intended application. Nucleotide synthesis involves a trade-off among procedural simplicity, conversion, isolated yield, and final purity. Direct one-pot approaches prioritize operational simplicity and may provide higher overall yields by avoiding intermediate purification and isolation steps, whereas protecting-group-based routes may provide greater control over chemoselectivity and product composition. Selection of an appropriate method should therefore consider the required purity, available equipment, reagent cost, accessibility, reaction scale, total synthesis time, and downstream purification capacity.
The method is particularly relevant to research programs aimed at producing nucleotide pools from biosynthetically derived nucleoside precursors, for which chemical phosphorylation can provide an accessible downstream conversion step. It may also be useful for preparing individual dNTPs for enzymatic assays, nucleotide-processing studies, nucleic acid synthesis, and the preliminary evaluation of modified nucleoside substrates. Furthermore, with substrate-specific optimization, the reaction sequence may be extended to the remaining canonical 2′-deoxyribonucleosides, providing a modular framework for the preparation of complete dNTP sets.
披露
The authors declare that there are no conflicts of interest related to the present work.
致谢
The authors thank the ”Sistema General de Regalías” which funded the project: “Producción de nucleótidos a partir de biomasa residual de la agroindustria para diagnóstico por biología molecular en el departamento de Santander” code BPIN 2021000100331.
材料
| 姓名 | 公司 | 目录编号 | 评论 |
|---|---|---|---|
| 2'-deoxyadenosine | Sigma Aldrich | D8668 | Nucleoside precursor |
| 2'-deoxyadenosine triphosphate | Sigma Aldrich | D6500 | Used as a standard in chromatographic analysis |
| Bis(tributylammonium) pyrophosphate salt | N/A | N/A | This reagent is synthesized in the laboratory. |
| Dichloromethane | Honeywell | 300-4L | |
| HPLC | Shimadzu | N/A | Reference Nexera XS inert |
| Hypersil GOLD C18 reversed-phase HPLC column | Thermo Fisher Scientific | 25005-154630 | C18 reversed-phase column |
| Phosphoryl chloride (POCl3) | Merck/Sigma Aldrich | 822339 | |
| Tributylamine (Bu3N) | Thermo Fisher Scientific | 139321000 | |
| Triethylammonium bicarbonate solution | N/A | N/A | This reagent is synthesized in the laboratory. |
| Trimethyl phosphate (Me3PO4) | Thermo Fisher Scientific | 157975000 | |
| Xelsius reactor | Lab Tech | 8053 100 002 | compact and fast reactor with 10 completely individually tempered and stirred reactor cells in the temperature range from -20°C to +150°C. |
参考文献
- Westarp S, Kaspar F, Neubauer P, Kurreck A. Industrial potential of the enzymatic synthesis of nucleoside analogs: existing challenges and perspectives. Curr Opin Biotechnol. 2022;78:102829.
- Ludwig J. A new route to nucleoside 5'-triphosphates. Acta Biochim Biophys Acad Sci Hung. 1981;16:131-3.
- Ludwig J, Eckstein F. Rapid and efficient synthesis of nucleoside 5'-O-(1-thiotriphosphates), 5'-triphosphates and 2',3'-cyclophosphorothioates using 2-chloro-4H-1,3,2-benzodioxaphosphorin-4-one. J Org Chem. 1989;54(3):631-5.
- Rachwalak M, Romanowska J, Sobkowski M, Stawinski J. Nucleoside di- and triphosphates as a new generation of anti-HIV pronucleotides. Chemical and biological aspects. Appl Sci. 2021;11:2248.
- Mohamady S, Taylor SD. Synthesis of nucleoside triphosphates from 2′,3′-protected nucleosides using trimetaphosphate. Org Lett. 2016;18(3):580-3.
- Vaghefi MM. Nucleoside triphosphates and their analogs: Chemistry, biotechnology, and biological applications. CRC Press; Boca Raton, FL; 2005.
- Gillerman I, Fischer B. An improved one-pot synthesis of nucleoside 5′-triphosphate analogues. Nucleosides Nucleotides Nucleic Acids. 2010;29(3):245-56.
- Hollenstein M. Nucleoside triphosphates — building blocks for the modification of nucleic acids. Molecules. 2012;17(11):13569-91.
- Taktakishvili M, Nair V. A new method for the phosphorylation of nucleosides. Tetrahedron Lett. 2000;41(37):7173-6.
- Liao JY et al. P(V) reagents for the scalable synthesis of natural and modified nucleoside triphosphates. J Am Chem Soc. 2019;141(34):13286-9.
- Johnson DC 2nd, Widlanski TS. Overview of the synthesis of nucleoside phosphates and polyphosphates. Curr Protoc Nucleic Acid Chem. 2004;Chapter 13:Unit 13.1.
重印与许可
本文已发表
视频即将推出