This protocol establishes a stratified genomic workflow for Salmonella serotype prediction, utilizing MLST or SeqSero2S for routine surveillance, SISTR with core genome MLST for outbreak detection, and SeqSero2S for rapid screening. The protocol has been validated through a multicenter analysis to enhance surveillance in China.
Method Article
A Stratified Genomic Framework for Salmonella Serotype Prediction: Evaluation of MLST, SeqSero, SeqSero2, SeqSero2S, and SISTR in Southwest China
* These authors contributed equally
In This Article
Summary
Abstract
Accurate serotyping of Salmonella is essential for effective surveillance and outbreak investigation, as serotype diversity directly impacts pathogenicity and public health risk assessment. However, conventional slide agglutination methods are limited by poor reproducibility, labor intensity, and high costs, which hinder their application in high-throughput monitoring programs. To address these limitations, we developed and validated a genomic workflow integrating Multilocus Sequence Typing (MLST), the Salmonella In Silico Typing Resource (SISTR), SeqSero, SeqSero2, and SeqSero2S for serotype prediction using whole-genome sequencing data. This protocol was evaluated through a multicenter analysis of 315 Salmonella isolates collected from food and human sources in Southwest China. The findings of this study demonstrated significantly higher concordance among genomic approaches (up to 100%/99.1%/90.1% in the training set and 100%/97.1%/93.1% in the validation set for SISTR/MLST/SeqSero2S, respectively) compared to traditional serotyping. The workflow includes recommendations for selecting appropriate prediction methods based on surveillance context, emphasizing MLST and SeqSero2S for routine monitoring, SeqSero2S for rapid screening, and SISTR with core genome MLST for outbreak investigations. This approach facilitates the integration of genomic serotyping into public health practice, reducing reliance on traditional serology and improving reproducibility and scalability in Salmonella monitoring programs.
Introduction
Salmonella is among the most significant foodborne pathogens globally, causing substantial morbidity and mortality1,2. In China, it remains a leading cause of foodborne illnesses, with a limited number of serotypes accounting for the majority of infections3. Accurate serotype identification is crucial for surveillance and outbreak investigations, as serotype diversity is closely correlated with pathogenicity and public health risk. This is well-recognized in common serotypes such as Salmonella enteritidis and Salmonella typhimurium, but some less studied serotypes, such as Salmonella Panama, are also a significant cause of global spread and invasive disease4, highlighting the general need for robust serotyping capabilities. Traditional serotyping methods, long recognized as the gold standard, are a time-consuming and high-cost process that requires over 150 antisera5. They are further constrained by poor reproducibility6, labor-intensive workflows7, and inherent variability due to subjective interpretation of agglutination reactions8, thus making them inadequate for high-throughput monitoring programs9.
The advent of whole-genome sequencing (WGS) has transformed pathogen characterization by enabling rapid, high-resolution serotype prediction10,11,12. Several in silico tools have been developed, including Multilocus Sequence Typing (MLST), SeqSero13, SeqSero214, SeqSero2S15, and the Salmonella In Silico Typing Resource (SISTR)7,15. While MLST offers standardized, long-term epidemiological tracking16, SeqSero2 and SeqSero2S allow direct analysis from raw reads and facilitate high-throughput screening of common serotypes14,15. Conversely, SISTR requires assembled genomes but integrates antigen gene detection with core genome multilocus sequence typing (cgMLST), providing comprehensive resolution for outbreak investigations17. While WGS-based Salmonella characterization is increasingly adopted as primary evidence in research16,18,19, its application in large-scale surveillance remains limited, a gap partly attributed to unaddressed practical considerations that influence method suitability. A global survey revealed that only 8% of participating laboratories use WGS for routine surveillance, with analytical complexity being a major barrier20. For instance, reliable serotype prediction hinges on adequate sequencing depth (typically >75× for cost-effectiveness, ideally ~200× for high-quality analysis) and high-quality genome assemblies (N50 of contigs > 30 kbp as a cost-effective minimum, preferably >100 kbp), particularly for assembly-dependent tools like SISTR21,22,23. Additionally, tool selection must align with a laboratory's computational resources and expertise: open-source tools such as SeqSero2S offer faster alternatives for rapid screening by analyzing raw reads and bypassing computationally intensive assembly15, while laboratories with limited bioinformatics capacity may struggle with data analysis and storage. Consequently, insufficient data exist to determine whether serological testing can be reduced in large-scale surveillance20,24, thereby highlighting the need for resource-adaptive approaches. Notably, following the COVID-19 pandemic, there is a substantial amount of idle sequencing instrumentation and qPCR across various levels of healthcare institutions, presenting an opportunity to repurpose this capacity for genomic surveillance.
To address this gap, we conducted a multicenter evaluation of the above-mentioned genomic serotyping tools using 315 Salmonella isolates collected from food and human sources in Southwest China25,26. Based on these findings, we developed a stratified workflow that accounts for varying laboratory resources and surveillance needs, providing clear criteria for tool selection in different contexts. This framework prioritizes SeqSero2S for rapid screening and SISTR combined with cgMLST for outbreak investigations, thereby facilitating the adoption of genomic approaches and reducing dependence on traditional serotyping methods. Here, we describe each step of the workflow in detail, including isolate identification, sequencing, data analysis, and criteria for selecting prediction tools in different surveillance contexts.
Protocol
The collection and handling of human fecal samples were conducted under approval from the Ethics Committee of West China Fourth Hospital, Sichuan University (Protocol Number: HXSY-EC-2022093), in accordance with the Helsinki Declaration and Good Clinical Practice. Informed consent was waived in accordance with national public health regulations.
NOTE: All procedures involving live Salmonella cultures and human fecal samples must be conducted in a Biosafety Level 2 (BSL-2) laboratory by personnel trained in standard microbiological practices and biosafety procedures. Wear appropriate personal protective equipment (PPE) at all times, including a lab coat, gloves, and safety goggles. Disinfect and change contaminated gloves regularly. Decontaminate all work surfaces before and after experiments with an appropriate disinfectant (e.g., 70% ethanol, 1% sodium hypochlorite). Decontaminate all liquid and solid waste containing biological materials by autoclaving (e.g., at 121 °C for at least 30 min) before disposal. Avoid the generation of aerosols and splashes. Perform all procedures that may create aerosols (e.g., centrifugation, vigorous shaking) inside a certified biological safety cabinet.
1. Food sample collection and strain identification
- Collect food samples from retail markets or processing facilities. Isolate Salmonella from food samples according to national food safety standard GB 4789.4 and the Foodborne Disease Surveillance Program's confidentiality protocols.
- Pre-enrichment:
- Weigh exactly 25 g of each food sample. Add 225 mL of Buffered Peptone Water (BPW) to the sample (1:10 dilution ratio) and vortex for 30 s to ensure full homogenization and submersion.
- Incubate the sample in BPW at 36°C ± 1°C for 8-18 h in a shaking incubator set to 150 rpm.
NOTE: Pre-enrichment in non-selective broth significantly increases the pathogen load. Handle all subsequent materials (containers, pipettes, etc.) as highly infectious.
- Selective enrichment: After pre-enrichment, transfer 1 mL of the sample to Tetrathionate Broth (TTB). Incubate the TTB at 42 °C for 18-24 h without shaking.
- Isolation: After incubation, streak the enriched culture onto the following selective agar plates: Hektoen Enteric (HE) agar, Xylose Lysine Deoxycholate (XLD) agar, and CHROMagar Salmonella. Incubate the plates at 36 °C for 18-24 h.
NOTE: When streaking plates, be cautious to avoid creating aerosols and cross-contamination. On HE agar, presumptive Salmonella colonies typically appear as blue-green to blue in color, with or without black centers. The surrounding agar may take on a similar greenish hue. On XLD agar, presumptive Salmonella colonies are typically red with black centers. On CHROMagar, presumptive Salmonella colonies are characterized by their magenta to mauve color, as specified by the manufacturer. On each selective plate, expect well-isolated colonies after 18-24 h of incubation. Overgrown plates (>300 colonies) may require dilution and re-streaking. - Identification: Examine the plates for suspected Salmonella colonies. Identify suspected strains using an automated identification system with a Gram-negative (GN) identification card, following the manufacturer's instructions for analysis.
NOTE: Ensure the lot numbers of the GN cards are valid and within the expiration date.- Prepare a bacterial suspension from a pure colony to the specified turbidity (0.50-0.63 McFarland standard) using 0.45% saline solution.
- Load the suspension into the GN card and run the test. Ensure the system is set to automatically incubate the card at 35.0 °C ± 2.0 °C and monitor biochemical reactions for up to 8 h.
- Accept a result of Salmonella enterica ssp. enterica with a ≥ 95% probability and Excellent confidence rating.
- Serotyping: Pick single colonies from the Swarm agar plate after overnight incubation at 37 °C. Perform slide agglutination tests to detect O-antigens and H-antigens. Use commercially available Salmonella antiserum kits (see Table of Materials) for testing.
2. Human sample collection and strain identification
NOTE: For the isolation of Salmonella from human fecal samples, follow the Foodborne Disease Surveillance Program's confidentiality protocols.
- Collection: Collect approximately 2 g of freshly voided human fecal samples. Place the fecal samples in clean, dry, wide-mouth containers. Transport the fecal samples to the laboratory within 1 h using insulated transport boxes to maintain sample integrity.
CAUTION: Human fecal samples may contain multiple pathogens beyond Salmonella. Handle all samples as potentially infectious and adhere to standard precautions for biohazard material handling. - Inoculation: Inoculate the collected fecal samples directly onto the following selective agar plates: CHROMagar Salmonella, Salmonella-XLD agar, and Shigella (SS) agar.
NOTE: Refer to step 1.4 for the characteristic colony morphology on CHROMagar Salmonella and XLD agar. On SS agar, presumptive Salmonella colonies typically appear as colorless, opaque, or tan with black centers. The agar itself remains relatively unchanged. - Identification: Identify suspected Salmonella colonies as described in step 1.5.
- Serotyping: Perform serotyping by slide agglutination as described in step 1.6.
3. Library preparation and whole-genome sequencing
- Bacterial culture: Streak Salmonella on LB agar, incubate at 37 °C for 12 h. Pick a colony, inoculate into 200 mL LB broth, and shake at 150 rpm (37 °C, 12 h). Centrifuge at 12,000 × g for 10 min to harvest cells.
NOTE: Perform all centrifugation steps with sealed rotors or safety cups to prevent aerosol generation. - DNA extraction: Extract genomic DNA using a commercial DNA extraction kit (see Table of Materials), following the manufacturer's protocol.
CAUTION: The reagents in commercial DNA extraction kits (e.g., lysis buffers, proteinase K) may be harmful if inhaled or upon contact with skin and eyes. Perform the extraction in a fume hood or biological safety cabinet if volatile reagents are used, and wear appropriate PPE. - DNA quality assessment: Purify genomic DNA and quantify using a fluorometer with the dsDNA High-Sensitivity Assay Kit. Verify DNA integrity by electrophoresis on a 1% agarose gel at 100 V for 45 min; high-quality DNA appears as a single compact band >20 kbp with minimal smearing27.
NOTE: (Critical step) High-quality, high-molecular-weight genomic DNA (with an A260/A280 ratio of ~1.8-2.0) is crucial for successful library preparation and sequencing.
CAUTION: The nucleic acid staining dyes used in agarose gel electrophoresis (e.g., GelRed, Ethidium Bromide) are mutagenic. Always wear gloves when handling gels and staining solutions, and dispose of them according to institutional hazardous waste regulations. - DNA fragmentation: Fragment approximately 150 ng of qualified genomic DNA to an average size of ~350 bp using a focused-ultrasonication system according to the manufacturer's protocol.
- Construct the sequencing library from the sheared DNA fragments using a commercial library preparation kit (see Table of Materials). Perform end-repair and 5'-phosphorylation of the sheared DNA fragments. Then, add a single 'A' base to the 3' ends, and ligate the indexed sequencing adapters.
- Amplify and enrich the adapter-ligated DNA fragments via a limited-cycle PCR program as specified in the kit.
- Qualify and quantify the final library using methods such as fluorometry and capillary electrophoresis.
- Perform paired-end sequencing (2 × 150 bp) on a sequencing platform following the standard operational procedures.
4. Data analysis
- Quality control of raw sequencing data: Perform initial quality control on the raw FASTQ files using fastp (version 0.23.0) with default parameters. The goal is to remove adapter sequences and low-quality reads, thereby generating clean FASTQ data for downstream analysis.
- Assemble the high-quality, filtered reads into draft genomes using SPAdes (version 3.15.0) with the careful option to reduce mismatches and short indels26.
- Utilize the genome sequences (FASTQ /FASTA files) as input for the various serotype prediction.
- For MLST, import the FASTQ file into BioNumerics software (version 7.6).
- In the Import Template window, click Edit parsing under advanced options. Set the Data parsing string and Data decoration to [DATA]. Apply the regular expressions DATA]_* and [DATA] for filename parsing. Save the template, select it, and click Next then Finish. Confirm submission in the Submit Task window to upload data. The CEStoreUploader tool will automatically open to transfer WGS raw data to the calculation engine.
- Select the genome entry after upload. Navigate to WGS Tools and select "TraNetGenotype" for automatic ST prediction. In the TraNetGenotype dialog box, select Salmonella as the strain type, select Serotype Analysis, Virulence Analysis, and Antimicrobial Susceptibility Analysis, and click Yes.
- For SISTR (version 1.1.3; https://github.com/phac-nml/sistr_cmd; Parameters: all parameters are default), open a terminal and navigate to the directory containing the assembled genome file assembly.fasta.
- Run the following command to perform serovar prediction and generate a comprehensive report in CSV format: sistr --qc --alleles-output allele-results.json --cgmlst-profiles cgmlst-profiles.csv -f tab -o sistr-output.tab -i assembly.fasta. This command generates a report containing the predicted serovar.
NOTE: -qc: Performs quality control checks on the assembly. -i assembly.fasta: Specifies the input FASTA file. -f csv: Defines the output format as CSV. -o sistr_results: Sets the base name for the output file (e.g., sistr_results.csv). -p 4: Uses 4 CPU threads for the analysis to speed up computation. --alleles-output: saves allele sequences and information to a JSON file. --cgmlst-profiles: exports cgMLST allele profiles to a CSV file.
- Run the following command to perform serovar prediction and generate a comprehensive report in CSV format: sistr --qc --alleles-output allele-results.json --cgmlst-profiles cgmlst-profiles.csv -f tab -o sistr-output.tab -i assembly.fasta. This command generates a report containing the predicted serovar.
- For SeqSero (version 1.0; https://github.com/denglab/SeqSero; Parameters: all parameters are default), navigate to the directory containing the input data file. Run the following command with default parameters for analysis: python SeqSero.py -m 4 -i assembly.fasta. The tool creates an output directory containing the file Seqsero_result.txt with the predicted serotype.
NOTE: -m 4 specifies input type as genome assembly. -i defines the input FASTA file. - For SeqSero2 (version 1.3.1; https://github.com/denglab/SeqSero2; Parameters: all parameters are default). Run the following command: python3 SeqSero2_package.py -m k -t 4 -i assembly.fasta. The tool outputs the predicted serotype directly to the terminal screen upon completion. The output provides the predicted serotype.
NOTE: -m k: specifies k-mer-based workflow for rapid serotype prediction using unique k-mers of serotype determinants. -t 4: indicates input type as assembled sequences. - For SeqSero2S (version 1.1.1; https://github.com/denglab/SeqSero2S; Parameters: all parameters are default): Run it using the provided script: python SeqSero2S.py -m k -t 4 -i assembly.fasta. The output provides the predicted serotype.
NOTE: -m k enables k-mer-based analysis of raw reads or genome assemblies. -t 4 specifies assembled sequence input.
- For MLST, import the FASTQ file into BioNumerics software (version 7.6).
5. Critical parameters and practical considerations
NOTE: The following considerations are crucial for the successful implementation of this genomic workflow, especially in resource-limited settings.
- Verify that the raw sequencing data achieves a sufficient depth of coverage. A sequencing depth of approximately 100x is typically sufficient for high-quality analysis.
NOTE: For laboratories prioritizing cost-effectiveness, a minimum sequencing depth of 75x is recommended as a practical lower threshold for reliable serotype prediction using MLST or SeqSero221. - Confirm that the quality of the raw reads is high, with a Q30 score (base call accuracy of 99.9%) typically above 90%.
- Assess the quality of the de novo genome assembly before proceeding with serotype prediction tools that require assembled contigs. Use the N50 statistic as a key metric.
NOTE: An N50 of at least 100 kbp is recommended22, while an N50 greater than 30 kbp should be regarded as the minimum requirement for reliable analysis23. - Check the completeness of the assembly by ensuring a read mapping rate of at least ≥98% when the filtered reads are mapped back to the assembled genome.
6. Recommended serotype prediction protocol for high-throughput salmonella surveillance
NOTE: Select serotype prediction methods based on surveillance context, required accuracy, and available laboratory resources. For high-throughput Salmonella surveillance, follow the protocol below:
- Preliminary Identification: Conduct initial screening in accordance with GB 4789.4 to confirm isolates as Salmonella spp.
NOTE: At this stage, there is no need to identify the specific serotype of the isolates. - Routine surveillance (High-Throughput mode): For routine surveillance, since BioNumerics is no longer updated, use open source MLST28, SeqSero2S, or other validated, user-friendly genomic analysis platforms that receive ongoing technical support and updates.
NOTE: For MLST: The 7-gene scheme offers a standardized, portable, and phylogenetically informative profile (Sequence Type, ST) that is ideal for long-term epidemiology and global strain comparison. Its simplicity ensures fast analysis and minimal computational burden. For SeqSero2S: It provides the dual advantage of rapid serotype prediction and integrated MLST results in a single run. It can analyze raw sequencing reads directly, bypassing genome assembly to save time and resources. Even with assembled FASTA data, it is faster than tools like SISTR, as it performs serotype prediction independently without relying on computational analysis. - Rapid screening (Time-sensitive settings): For rapid results or in laboratories with limited computational capacity, use SeqSero2S for preliminary serotype prediction.
NOTE: When performing SeqSero2S predictions, it is basically recommended to interpret "I_4,[5],12:i:-" as Salmonella Typhimurium. - Outbreak investigation (High-accuracy mode): For outbreak tracing requiring high resolution, use SISTR, which uses cgMLST to supplement serotype prediction and to provide high-resolution cluster analysis for source tracing.
- Quality control and inconsistent results confirmation:
- For common sequencing type strains, perform random verification. For non-common ones, use a verification ratio of 20%. In cases of inconsistent serotyping results, refer isolates to a higher-level or reference laboratory for a definitive result.
- For confirmatory analysis, employ multiple validated methods including: repeated genomic analysis with at least two different in silico prediction tools; repeated in silico prediction with at least two different tools; phylogenetic contextualization using high-resolution WGS-based methods (e.g., cgMLST or whole-genome SNP analysis)29.
- Assign the final serotype based on consensus from this comprehensive evidence.
NOTE: The recommended 20% verification ratio for non-common serotypes is based on the observation that genomic tools maintain high concordance (93.3% for SISTR vs. MLST, 95.0% for SISTR vs. SeqSero2S) even among non-common serotypes (Table 1). This threshold is cost-effective and provides a sufficient safety margin to detect the low frequency of discrepancies among genomic tools in routine surveillance.
7. Statistical analysis
- Perform data analysis using R software (version 4.1.2).
- Present categorical variables as frequencies with percentages.
- Compare groups using χ2 tests for categorical data30, consider a P-value <0.05 as statistically significant.
Results
The protocol was validated using 315 Salmonella isolates from Southwest China, with 213 isolates (2/3) serving as the training set to compare five WGS-based serotype prediction methods (MLST, SISTR, SeqSero, SeqSero2, and SeqSero2S) against conventional serotyping results, while the remaining 102 isolates (1/3) were used as a validation set to assess MLST, SISTR, and SeqSero2S performance.
Training set analysis (n = 213)
Reproducibility assessment: Conventional serotyping demonstrated limited concordance with any of the five WGS-based prediction methods, achieving only 60.6% agreement (129/213). In contrast, pairwise agreement among any two of the five WGS approaches reached 99.1% (211/213), and complete five-method consensus was observed in 82.2% of isolates (175/213). This substantial disparity (82.2% vs. 60.6%, P<0.001) underscores the superior reproducibility of genomic approaches compared to conventional serology (Figure 1).
Concordance ranking: Comparison among WGS prediction methods showed the following descending order of concordance rates: SISTR (100%, 213/213), MLST (98.1%, 209/213), SeqSero2S (90.1%, 192/213), and SeqSero2/SeqSero (both 83.1%, 177/213). While no significant difference existed between MLST and SISTR (P=0.123), both MLST and SISTR showed significantly higher concordance than SeqSero/SeqSero2/SeqSero2S (P<0.001). Furthermore, SeqSero2S demonstrated statistically superior performance relative to both SeqSero and SeqSero2 (P=0.033), establishing a clear performance gradient: SISTR> MLST> SeqSero2S>SeqSero2≈SeqSero (Figure 1, Supplementary File 1).
Validation set confirmation (n = 102)
Reproducibility assessment: Concordance between conventional serotyping and any WGS methods was 66.7% (68/102), while pairwise WGS agreement reached 100% (102/102), with five-method consensus in 88.2% (90/102), reaffirming the superior reproducibility of WGS-based methods (88.2% vs. 67.6%, P=0.004) (Figure 2).
Concordance ranking: WGS methods showed similar concordance: SISTR (100.0%, 102/102), MLST (97.1%, 99/102), SeqSero2S (93.1%, 95/102), and SeqSero2/SeqSero (both 91.2%,93/102). While MLST and SISTR showed no significant difference (P=0.246), only SISTR showed significantly higher concordance than SeqSero/SeqSero2/SeqSero2S (P<0.05). The same performance hierarchy was observed: SISTR> MLST> SeqSero2S>SeqSero2≈SeqSero (Figure 2, Supplementary File 1).
Total set evaluation (n = 315)
In the comprehensive evaluation of all 315 samples, SISTR demonstrated complete coverage (100%, 315/315). MLST and SeqSero2S also showed high concordance rates of 97.8% (308/315) and 91.1% (287/315), respectively. The superior coverage of genomic methods contrasted sharply with traditional serotyping (84.1% (265/315) vs. 62.9% (198/315), P<0.001), highlighting its limitations as a primary surveillance tool (Figure 3A).
To systematically evaluate method performance, we assessed pairwise concordance across all 315 isolates. As shown in the inter-method concordance heatmap (Figure 3B), genomic approaches exhibited exceptional mutual agreement, with consistency scores exceeding 0.89 among SISTR, MLST, and SeqSero2S. In stark contrast, traditional serotyping showed substantially lower concordance (0.49-0.57) with all genomic platforms, revealing a fundamental methodological divergence.
The consensus network analysis (Figure 3C) further confirmed these relationships, showing genomic methods forming a tightly interconnected core with SISTR and SeqSero2S as central hubs. The robust connections among genomic tools underscore their reliability, while the weak links connecting traditional serotyping visually emphasize its inconsistent performance.
For detailed serotype-level analysis, we used SISTR predictions as the benchmark due to its complete coverage and examined concordance with other methods in Table 1. Among the 15 major Salmonella serotypes (n≥5), MLST demonstrated perfect concordance (100%) with SISTR for 13/15 serotypes, while SeqSero2S achieved 100% concordance for 11/15 serotypes. Remarkably, conventional serotyping failed to achieve perfect concordance with SISTR for any of the 15 major serotypes (0/15), with concordance rates ranging from 0% to 95.0% (Table 1, Supplementary File 1).
Interpretation guidelines
For reliable prediction: Consistent with the high inter-method agreement observed among genomic tools, a consensus among SISTR, MLST, and Seqsero2S (concordance ≥90% in this study) provides a highly reliable serotype prediction.
For tool selection: The performance hierarchy (SISTR ≈ MLST > SeqSero2S) provides a data-driven basis for selecting tools based on required accuracy and resources, as detailed in the recommended workflow (section 6).
Suboptimal case
Genomic vs. traditional serotyping discrepancies: The low concordance (0.49-0.57) between genomic platforms and traditional serotyping reflects a fundamental methodological divergence. In cases of discrepancy, the consensus from genomic methods (especially SISTR and MLST) should be prioritized, as they demonstrate superior reproducibility and coverage.
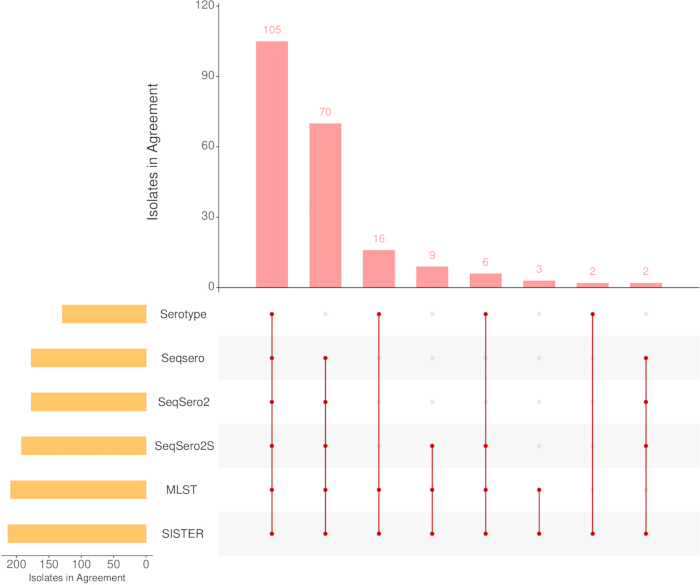
Figure 1: UpSet plot comparing traditional serotyping with five WGS-based prediction methods (MLST, SeqSero, SeqSero2, SeqSero2S, SISTR) in the test set (n = 213). This UpSet plot visualizes the agreement and discrepancies between conventional serotyping and the five whole-genome sequencing (WGS)-based prediction methods (MLST, SeqSero, SeqSero2, SeqSero2S, SISTR). The horizontal bar chart (left) shows the size of each set (the number of isolates in agreement assigned to each method). The vertical bar chart (top) shows the size of the intersections between methods (the number of isolates where predictions agreed). The dot and line matrix (bottom) indicates which methods are included in each intersection. In the test set (n = 213), significantly higher inter-method agreement among the five WGS-based approaches (top3: SISTR, MLST, SeqSero2S) compared to their concordance with traditional serotyping. Please click here to view a larger version of this figure.
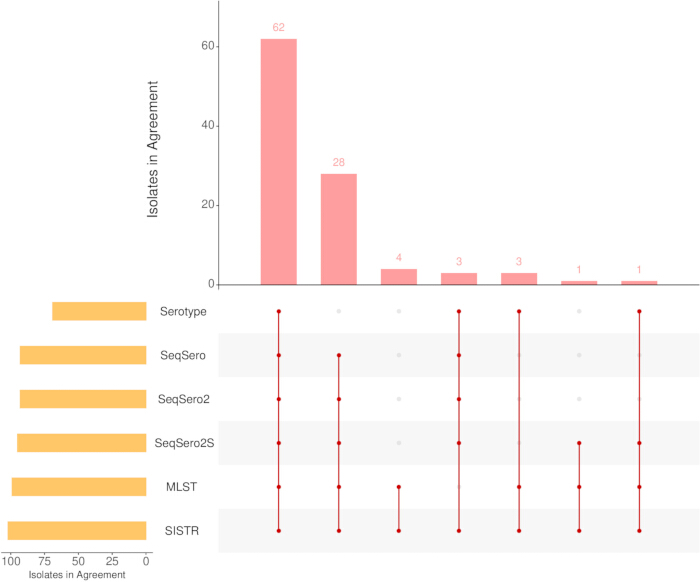
Figure 2: UpSet plot validating serotyping against five WGS-based methods (MLST, SeqSero, SeqSero2, SeqSero2S, SISTR) in the verification set (n = 102). This UpSet plot validates the findings from the training set. The results reaffirm the superior reproducibility of WGS-based methods, as indicated by the large intersection among SISTR, MLST, and SeqSero2S, compared to their smaller shared intersections with traditional serotyping. Please click here to view a larger version of this figure.
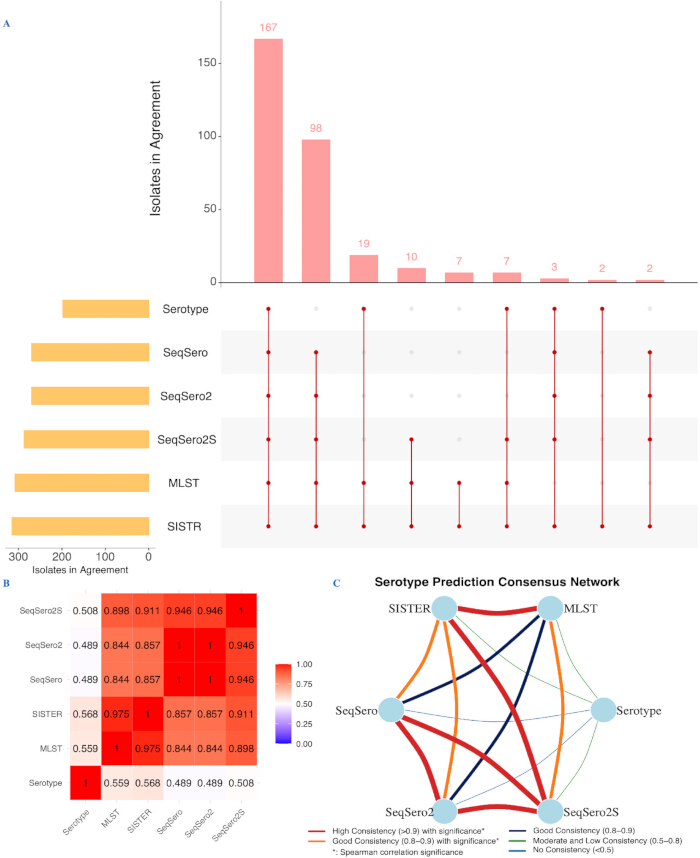
Figure 3: Comparison of serotype prediction concordance between traditional serotyping and five in silico genomic methods (n = 315). (A) UpSet plot of method agreement patterns. SISTR achieves complete coverage (315/315, 100%), followed by MLST (308/315, 97.8%), SeqSero2S (287/315, 91.1%), SeqSero2 (270/315, 85.7%), SeqSero (270/315, 85.7%), and traditional serotyping (198/315, 62.9%). (B) Heatmap of inter-method concordance. Genomic methods (MLST, SISTR, SeqSero2S) demonstrate high mutual agreement (>0.89), whereas traditional serotyping shows substantially lower concordance with these genomic methods (0.49-0.57). The strong concordance among genomic platforms supports their reliability for Salmonella surveillance. (C) Consensus network of serotyping approaches. This network visualization depicts concordance relationships among the six serotyping approaches, where edge thickness and color intensity represent the level of agreement. The circular layout reveals two distinct clusters: genomic methods form a tightly interconnected core with strong consistency (>0.84), while traditional serotyping remains peripherally connected with weaker links (0.49-0.57). SeqSero2S shows robust connections to all genomic methods (>0.90). Please click here to view a larger version of this figure.
Table 1: Concordance between SISTR-predicted serotypes and conventional serotyping/MLST/ SeqSero2S results for Salmonella strains (n = 315). Please click here to download this Table.
Supplementary File 1: Clean data. Please click here to download this File.
Discussion
This large-scale evaluation, encompassing 315 Salmonella isolates from Southwest China, demonstrates the superior reproducibility of WGS-based serotyping methods compared to conventional approaches (84.1% vs. 62.9% coverage, P<0.001). Our findings validate a comprehensive genomic workflow where critical initial steps of sample preparation and DNA quality control (sections 1-3) directly impact downstream reliability. The protocol emphasizes high-quality genome assemblies (preferably >100 kbp) for assembly-dependent tools (sections 4-5). The stratified implementation strategy allows laboratories to select appropriate methods based on their specific surveillance needs and computational resources, with SISTR recommended for definitive analysis (100% concordance across both training and validation sets), MLST (98.1%/97.1%)/SeqSero2S (90.1%/93.1%) for routine surveillance, and SeqSero2S for rapid screening (section 6). This framework effectively addresses practical laboratory constraints, as SeqSero2S provides a viable alternative for resource-limited settings, bypassing computationally intensive assembly steps while maintaining 91.1% coverage15.
We observed reduced concordance for certain serotypes, particularly Salmonella Typhimurium where SeqSero2S identified 23/51 isolates as I_4,[5],12:i:-, highlighting challenges in detecting monophasic variants31,32. When performing SeqSero2S predictions, it is basically recommended to interpret "I_4,[5],12:i:-" as Salmonella Typhimurium.
For verification and to resolve discrepancies, we systematically employ a multi-pronged approach that includes concordance checks across genomic tools and phylogenetic placement via cgMLST or wgSNP analysis (step 6.5). A majority of common Salmonella serovars are polyphyletic, meaning that the same serotype can arise from multiple, genetically distinct lineages29. This finding underscores a critical limitation of serotyping alone and highlights the necessity of phylogenetic contextualization in our confirmatory workflow to validate the serotype prediction when results conflict. In such cases, cgMLST or wgSNP analysis provides confirmatory evidence for the genomic prediction29.
The eventual sunsetting of BioNumerics evokes the poetic imagery of heroes fading away like spray in the eastward-flowing Long River. This platform served as one of the most important analytical software for many public health laboratories19,25,26,33, providing an integrated and user-friendly gateway into the world of whole-genome sequencing analysis. Its discontinuation, however, represents not an endpoint but a natural evolution, compelling the field to embrace more open, flexible, and sustainable solutions28,34. The transition of MLST from BioNumerics to open-source alternatives could ensure workflow continuity without compromising analytical performance.
This protocol represents a significant advancement over conventional serotyping, demonstrating superior reproducibility of genomic methods (84.1% vs. 62.9%, P<0.001) while addressing the limitations of traditional approaches, including subjective interpretation and technical variability7,8. The high concordance between SISTR, MLST, and SeqSero2S for the major and minor Salmonella serotypes supports transitioning to genomic-based surveillance, reducing reliance on labor-intensive serology. The stratified framework optimizes resource allocation by matching tool selection to specific surveillance contexts, making genomic approaches accessible to laboratories with varying capabilities.
As WGS becomes more accessible, this stratified framework effectively integrates genomics into public health practice. It balances efficiency with accuracy, reduces reliance on labor-intensive serology, and enables rapid responses to public health threats35. The findings of this study, along with evidence from other surveillance systems, demonstrate that transitioning to WGS-based serotyping is both technically feasible and strategically advantageous for strengthening enteric pathogen surveillance36. It is important to note that the framework validated here is based on current mainstream rule-based algorithms. While tools like SISTR and SeqSero2 are highly useful, their performance is dependent on the quality and diversity of reference databases, which can limit their accuracy for emerging or atypical serotypes. Looking ahead, emerging machine learning methods have demonstrated high accuracy in serotype prediction37,38. However, it is important to recognize that these machine learning approaches currently face their own set of challenges before they can be widely deployed in routine surveillance. A primary limitation is their dataset dependency; models trained on one specific dataset often show a noticeable drop in accuracy when applied to another39. Furthermore, the "black box" nature of many complex models can obscure the specific genomic evidence used for a prediction, which is a critical requirement for public health attribution and investigative tracing. In addition, the performance of these models is closely tied to the quality and breadth of their training data40. They may struggle with novel or highly divergent serotypes not represented in the training set, and their operational utility in real-time, high-throughput surveillance pipelines against established, optimized tools remain to be thoroughly validated. These data-driven approaches represent a promising new direction for the future development of serotype prediction tools, but further development is needed to enhance model robustness, interpretability, and generalizability for public health applications.
In addition, future technological advancements will further enhance the capabilities of genome monitoring. For example, a new frontier in genome assembly is based on the hybrid assembly of short and long reads, which will produce a more complete reference genome, providing a more solid foundation for precise serotype prediction and source tracing6,41.
This validated genomic workflow addresses a critical gap in Salmonella surveillance by providing clear, evidence-based guidelines for method selection across different laboratory contexts. By balancing accuracy requirements with practical constraints, the protocol enables efficient implementation of WGS-based serotyping, ultimately strengthening enteric pathogen surveillance capabilities and enhancing public health response to Salmonella infections.
Disclosures
The authors have no conflicts of interest to declare.
Acknowledgements
This work was supported by grants from Tibet and Sichuan Science and Technology Bureau Funding (XZ202301ZY0049G, 2024NSFSC0563 to H.Z. and 2024NSFSC0033 to X.P.), Sichuan Science and Technology Program (2024JDRC0032 to Y.X.), the Project of Institute of Health New Productivity (HN240302C to H.Z.), the Discipline Revitalization Project of Public Health Laboratory Sciences (2023SY-04 to H.Z.), West China School of Public Health/West China Fourth Hospital, Sichuan University.
Materials
| Name | Company | Catalog Number | Comments |
|---|---|---|---|
| BioNumerics (version 7.6) | Applied Maths | https://bionumerics.software.informer.com/7.6/ | MLST/serotype prediction/phylogenic analysis |
| CHROMagar Salmonella | Becton, Dickinson and Company | 214983 | Selective Salmonella chromogenic medium |
| Fastp (version 0.23.0) | Open-source | https://github.com/OpenGene/fastp | Read quality control |
| Illumina NovaSeq 6000 Sequencing System | Illumina | 20012850 | Whole-genome sequencing platform |
| M220 Focused-ultrasonicator | Covaris | 500295 | DNA shearing |
| Magbeads Fast DNA Kit for Soil | MP Biomedicals | MP116560200 | Genomic DNA extraction kit |
| Microsoft Excel 2016 | Microsoft | KB5002794 | Data management |
| NEXTFLEX Rapid DNA-Seq Kit | Revvity | 50-255-1124 | Sequencing library preparation |
| Qubit 2.0 Fluorometer | Thermo Fisher Scientific | Q32866 | DNA quantification |
| Qubit dsDNA High-Sensitivity Assay Kit | Thermo Fisher Scientific | Q32851 | DNA quantification |
| R (version 4.1.2) | R Foundation | https://www.r-project.org/ | Statistical computing |
| Salmonella Antiserum Kit | Statens Serum Institute | 60898 | O/H-antigen detection |
| SeqSero | Open-source | https://github.com/denglab/SeqSero | Serotype prediction |
| SeqSero2 | Open-source | https://github.com/denglab/SeqSero2 | Serotype prediction |
| SeqSero2S | Open-source | https://github.com/denglab/SeqSero2s | Serotype prediction |
| SISTR | Open-source | https://github.com/phac-nml/sistr_cmd | Serotype prediction |
| SPAdes (version 3.15.0) | Open-source | https://github.com/ablab/spades | Genome assembly |
| Swarm Agar | BD Biosciences | 211519 | Used for serotyping incubation |
| TBS-380 Fluorometer | Turner BioSystems | P/N 3800-003 | DNA quantification |
| VITEK 2 Compact System | BioMérieux | 95061-768 | Automated microbial identification system |
References
- GBD 2017 Non-Typhoidal Salmonella Invasive Disease Collaborators. The global burden of non-typhoidal Salmonella invasive disease: a systematic analysis for the Global Burden of Disease Study 2017. Lancet Infect Dis. 19 (12), 1312-1324 (2019).
- Selim, S. A. A comprehensive review of antibiotic susceptibility patterns in strains isolated from food. J Pure Appl Microbio. 19 (2), 818-833 (2025).
- Shen, Y., et al. Genomic investigation of Salmonella enterica serovar Welikade from a pediatric diarrhea case first time in Shanghai, China. BMC Genomics. 25 (1), 604(2024).
- Pulford, C. V., et al. Global diversity and evolution of Salmonella enterica serovar Panama: a genomic epidemiology study. Lancet Microbe. 6 (9), 101150(2025).
- Hong, H., et al. Genetic characterization and in silico serotyping of 62 Salmonella enterica isolated from Korean poultry operations. BMC Genomics. 26 (1), 166(2025).
- Kitchens, S. R., Wang, C., Price, S. B. Bridging classical methodologies in Salmonella investigation with modern technologies: a comprehensive review. Microorganisms. 12 (11), 2249(2024).
- Uelze, L., et al. Performance and accuracy of four open-source tools for in silico serotyping of Salmonella spp. based on whole-genome short-read sequencing data. Appl Environ Microbiol. 86 (5), e02265-e02319 (2020).
- Wang, B. X., et al. High-throughput fitness experiments reveal specific vulnerabilities of human-adapted Salmonella during stress and infection. Nat Genet. 56 (6), 1288-1299 (2024).
- Kumar, S., Kumar, Y., Kumar, G., Kumar, G., Kasana, D. A highly drug-resistant Salmonella enterica serovar Weltevreden of human origin from India and detection of its virulence factors. Indian J Microbiol. 65 (3), 1384-1394 (2025).
- Lu, Y., et al. Advances in whole genome sequencing: methods, tools, and applications in population genomics. Int J Mol Sci. 26 (1), 372(2025).
- Mukherjee, S., et al. Increasing frequencies of antibiotic resistant non-typhoidal Salmonella infections in Michigan and risk factors for disease. Front Med (Lausanne). 6, 250(2019).
- Yachison, C. A., et al. The validation and implications of using whole genome sequencing as a replacement for traditional serotyping for a national Salmonella reference laboratory. Front Microbiol. 8, 1044(2017).
- Zhang, S., et al. Salmonella serotype determination utilizing high-throughput genome sequencing data. J Clin Microbiol. 53 (5), 1685-1692 (2015).
- Zhang, S., et al. SeqSero2: rapid and improved Salmonella serotype determination using whole-genome sequencing data. Appl Environ Microbiol. 85 (23), e01746-e01819 (2019).
- Deng, X., et al. Salmonella serotypes in the genomic era: simplified Salmonella serotype interpretation from DNA sequence data. Appl Environ Microbiol. 91 (3), e0260024(2025).
- Cui, Q., et al. Characterization and implications of the regionally prevalent ST8333 strains of Salmonella enterica serotype 4,[5],12:i: - China, 2017-2023. China CDC Wkly. 6 (47), 1232-1235 (2024).
- Yoshida, C. E., et al. The Salmonella In Silico Typing Resource (SISTR): an open web-accessible tool for rapidly typing and subtyping draft Salmonella genome assemblies. PLoS One. 11 (1), e0147101(2016).
- Ikeuchi, S., et al. Molecular epidemiological analysis of Salmonella Schwarzengrund isolated in Japan by newly developed multi-locus variable-number tandem repeat analysis method. Lwt. 207, 116593(2024).
- Kong, X., et al. Emergence of extensively drug-resistant Salmonella Kentucky ST198 in Southwest China. J Glob Antimicrob Resist. 43, 264-270 (2025).
- Davedow, T., et al. PulseNet international survey on the implementation of whole genome sequencing in low and middle-income countries for foodborne disease surveillance. Foodborne Pathog Dis. 19 (5), 332-340 (2022).
- Xian, Z., et al. Subtyping evaluation of Salmonella Enteritidis using single nucleotide polymorphism and core genome multilocus sequence typing with nanopore reads. Appl Environ Microbiol. 88 (15), e0078522(2022).
- Liu, C. C., Hsiao, W. W. L. Machine learning reveals the dynamic importance of accessory sequences for Salmonella outbreak clustering. mBio. 16 (3), e0265024(2025).
- Fourteenth external quality assessment for Salmonella typing. , European Centre for Disease Prevention and Control. ECDC, Stockholm. (2025).
- Gomes, E., et al. Advances in whole genome sequencing for foodborne pathogens: implications for clinical infectious disease surveillance and public health. Front Cell Infect Microbiol. 15, 1593219(2025).
- Zhou, L., et al. Antimicrobial resistance and genomic investigation of Salmonella isolated from retail foods in Guizhou, China. Front Microbiol. 15, 1345045(2024).
- Zuo, H., et al. Comparative genomic and antimicrobial resistance profiles of Salmonella strains isolated from pork and human sources in Sichuan, China. Front Microbiol. 16, 1515576(2025).
- Feehan, J. M., et al. Purification of high molecular weight genomic DNA from powdery mildew for long-read sequencing. J Vis Exp. (121), e55463(2017).
- Jolley, K. A., Bray, J. E., Maiden, M. C. J. Open-access bacterial population genomics: BIGSdb software, the PubMLST.org website and their applications. Wellcome Open Res. 3, 124(2018).
- Yang, L., et al. Phylogeny and divergence of the 100 most common Salmonella serovars available in the NCBI Pathogen Detection database. Front Microbiol. 16, 1547190(2025).
- Zuo, H., et al. High-altitude exposure decreases bone mineral density and its relationship with gut microbiota: results from the China multi-ethnic cohort (CMEC) study. Environ Res. 215 (Pt 2), 114206(2022).
- Arrieta-Gisasola, A., et al. Genotyping study of Salmonella 4,[5],12:i:- monophasic variant of serovar Typhimurium and characterization of the second-phase flagellar deletion by whole genome sequencing. Microorganisms. 8 (12), 2049(2020).
- De Sousa Violante, M., et al. Genomic diversity of Salmonella Typhimurium and its monophasic variant in pig and pork production in France. Microbiol Spectr. 12 (12), e0052624(2024).
- Kong, X., et al. Phenotypic and genotypic characterization of Salmonella Enteritidis isolated from two consecutive food-poisoning outbreaks in Sichuan, China. J Food Safety. 43 (1), e13015(2022).
- Hong, W., et al. Integrating serotyping, MLST, and phenotypic data: decoding the evolutionary drivers of Salmonella pathogenicity and drug resistance. Appl Environ Microbiol. 91 (10), e0151125(2025).
- Lin, A., et al. Targeted next-generation sequencing assay for direct detection and serotyping of Salmonella from enrichment. J Food Prot. 87 (4), 100256(2024).
- van den Berg, O. E., et al. Ongoing increase in autochthonous Salmonella Enteritidis infections, the Netherlands, June 2023 to June 2025. Euro Surveill. 30 (30), 2500536(2025).
- Li, X., Oladeinde, A., Rothrock, M. Jr, Chung, T. J., Ghazi Al Hakeem, W. Using core genome and machine learning for serovar prediction in Salmonella enterica subspecies I strains. FEMS Microbiol Lett. 372, fnaf040(2025).
- Krüger, G. I., et al. Adaptive signatures of emerging Salmonella serotypes in response to stressful conditions in the poultry industry. Lwt. 215, 117188(2025).
- Gou, J., et al. Microbial biomarkers and sex-associated gut microbiota characteristics of thyroid cancer. BMC Cancer. 25, 1688(2025).
- Meng, J., et al. Medical laboratory data-based models: opportunities, obstacles, and solutions. J Transl Med. 23 (1), 823(2025).
- Wu, X., et al. Evaluation of multiplex nanopore sequencing for Salmonella serotype prediction and antimicrobial resistance gene and virulence gene detection. Front Microbiol. 13, 1073057(2022).